
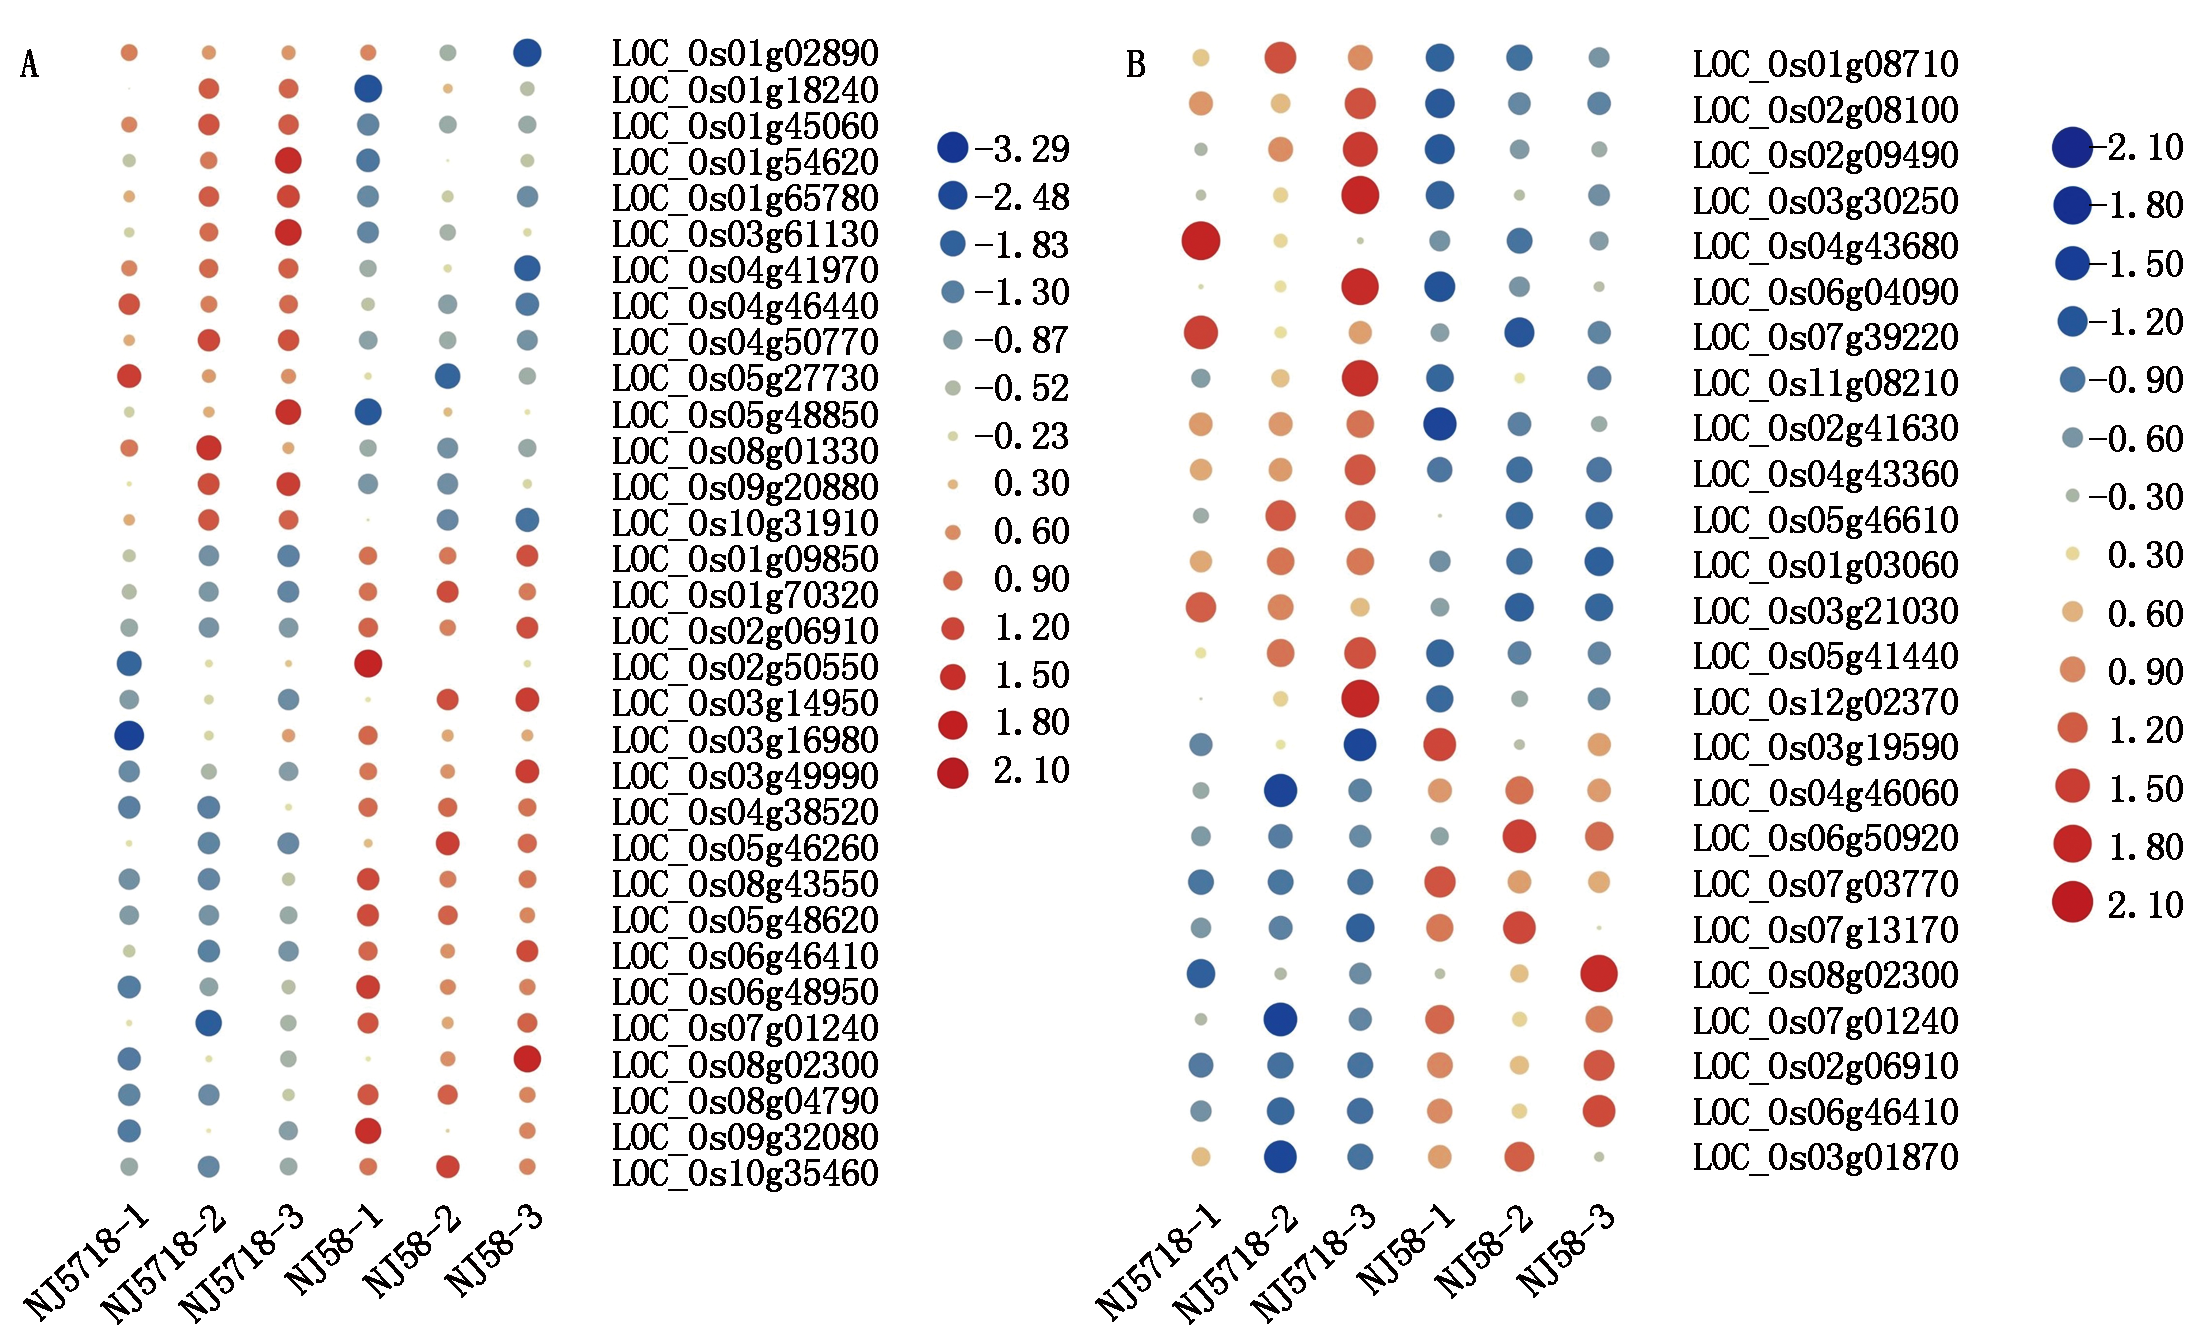
The super rice variety Nanjing 5718 has been designated as a leading variety by the Ministry of Agriculture and Rural Affairs and has demonstrated outstanding lodging resistance in large-scale production.Studying the stem characteristics of Nanjing 5718 and elucidating the molecular mechanisms underlying its lodging resistance can provide new insights for breeding lodging-resistant rice varieties.The lodging-resistant super rice Nanjing 5718 and two varieties,Nanjing 518 and Nanjing 58,with weak lodging resistance,were used as materials to compare agronomic traits,stem characteristics and their correlation with lodging resistance.The plant height and center of gravity of Nanjing 5718 were significantly lower than those of other varieties at 20 days after heading.The length of the second basal internode,stem diameter and stem wall thickness of the second and third basal internodes of Nanjing 5718 were significantly higher than other varieties at 20 days after heading.High content of cellulase and lignin in the stem was the important factor leading to strong lodging resistance.No significant differences in cellulase activity were observed in the second basal internodes among the three varieties.Compared to the other two varieties,only the pPLAⅢα,bHLH002,ILA1 and NAC5 among the 102 genes related to the cellulose and lignin biosynthetic pathways had missense mutations in their coding regions in Nanjing 5718.The relative expression levels of key cellulose biosynthesis-regulating genes,such as MYB61, SLR1,PIL13, PG3, ERF34,DPH1 and MYB7 were significantly different between Nanjing 5718 and Nanjing 58.Significant differences were also observed in the relative expression levels of some lignin metabolism genes between Nanjing 5718 and Nanjing 58:4CL3,RLM1,CYP98A4,SWN1,PAL1,and so on.The differential expression of those genes in basal internodes likely underlies the high cellulose and lignin content of stem and consequent strong lodging resistance in Nanjing 5718.In conclusion,the strong lodging resistance of the super rice variety Nanjing 5718 results from its increased internode diameter and stem wall thickness.The higher contents of cellulose and lignin in the internodes provide a structural basis for this enhanced lodging resistance.Variations in the expression of key genes involved in the cellulose and lignin biosynthesis pathways may be the primary factor contributing to the superior lodging resistance observed in Nanjing 5718.
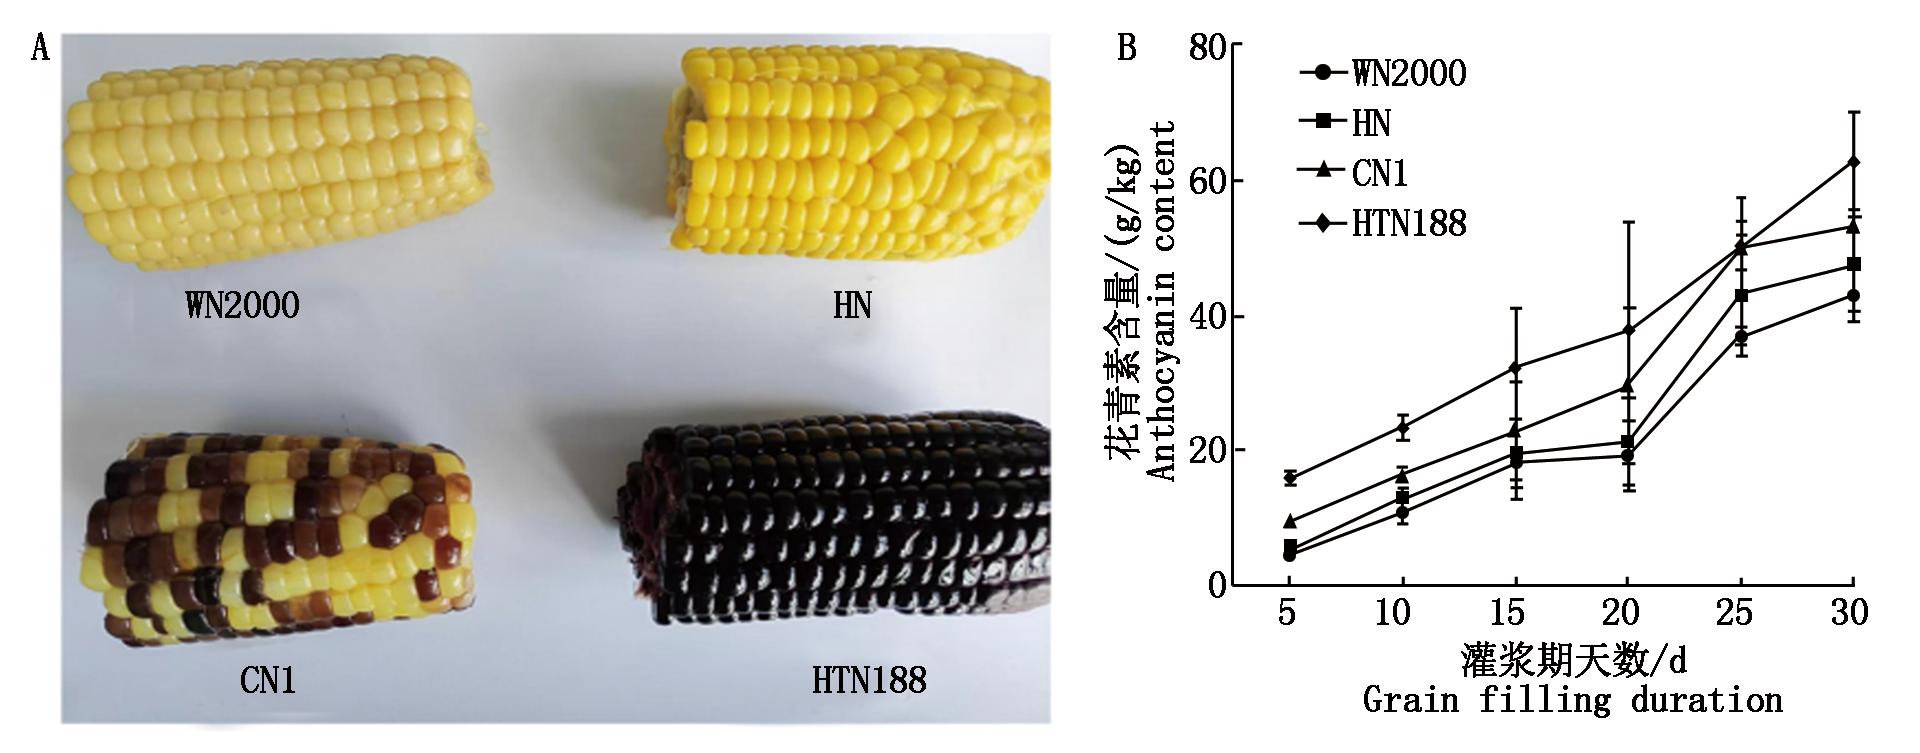
This study aimed to elucidate the compositional differences and molecular regulatory mechanisms of anthocyanins in kernels of fresh-eating maize with different colors by integrating metabolomic and transcriptomic analyses,thereby providing candidate genes and a theoretical basis for molecular breeding of colored fresh-eating maize.White WN2000,yellow HN,colored CN1,and black HTN188 were used as materials.Metabolomics was employed to identify anthocyanin-related metabolites and screen differential metabolites;transcriptome sequencing was conducted to obtain differentially expressed genes(DEGs).Metabolite gene associations were analyzed through GO annotation and KEGG pathway enrichment,and key structural genes and transcriptional regulators were mined within the anthocyanin biosynthetic pathway.The results showed that 49 anthocyanin-related metabolites were identified in total.Major anthocyanins,including cyanidin,peonidin,and pelargonidin,accumulated significantly in HTN188,indicating stronger pigment deposition.Transcriptome analysis identified 12 557 DEGs,among which structural genes such as F3H,DFR,and UFGT,as well as transcription factors including MYB and bHLH,were significantly up-regulated in varieties with high anthocyanin content.Correlation analysis revealed that the expression level of Zm00001d011438 was significantly and positively correlated with total anthocyanin content,suggesting that it may play a key role in kernel anthocyanin accumulation.Overall,the integrated metabolomic and transcriptomic analyses characterized differences in anthocyanin composition and related gene expression among differently colored fresh-eating maize kernels,and identified structural genes,transcription factors,and the candidate Zm00001d011438 associated with the anthocyanin biosynthetic pathway,providing references for subsequent functional validation and genetic improvement.
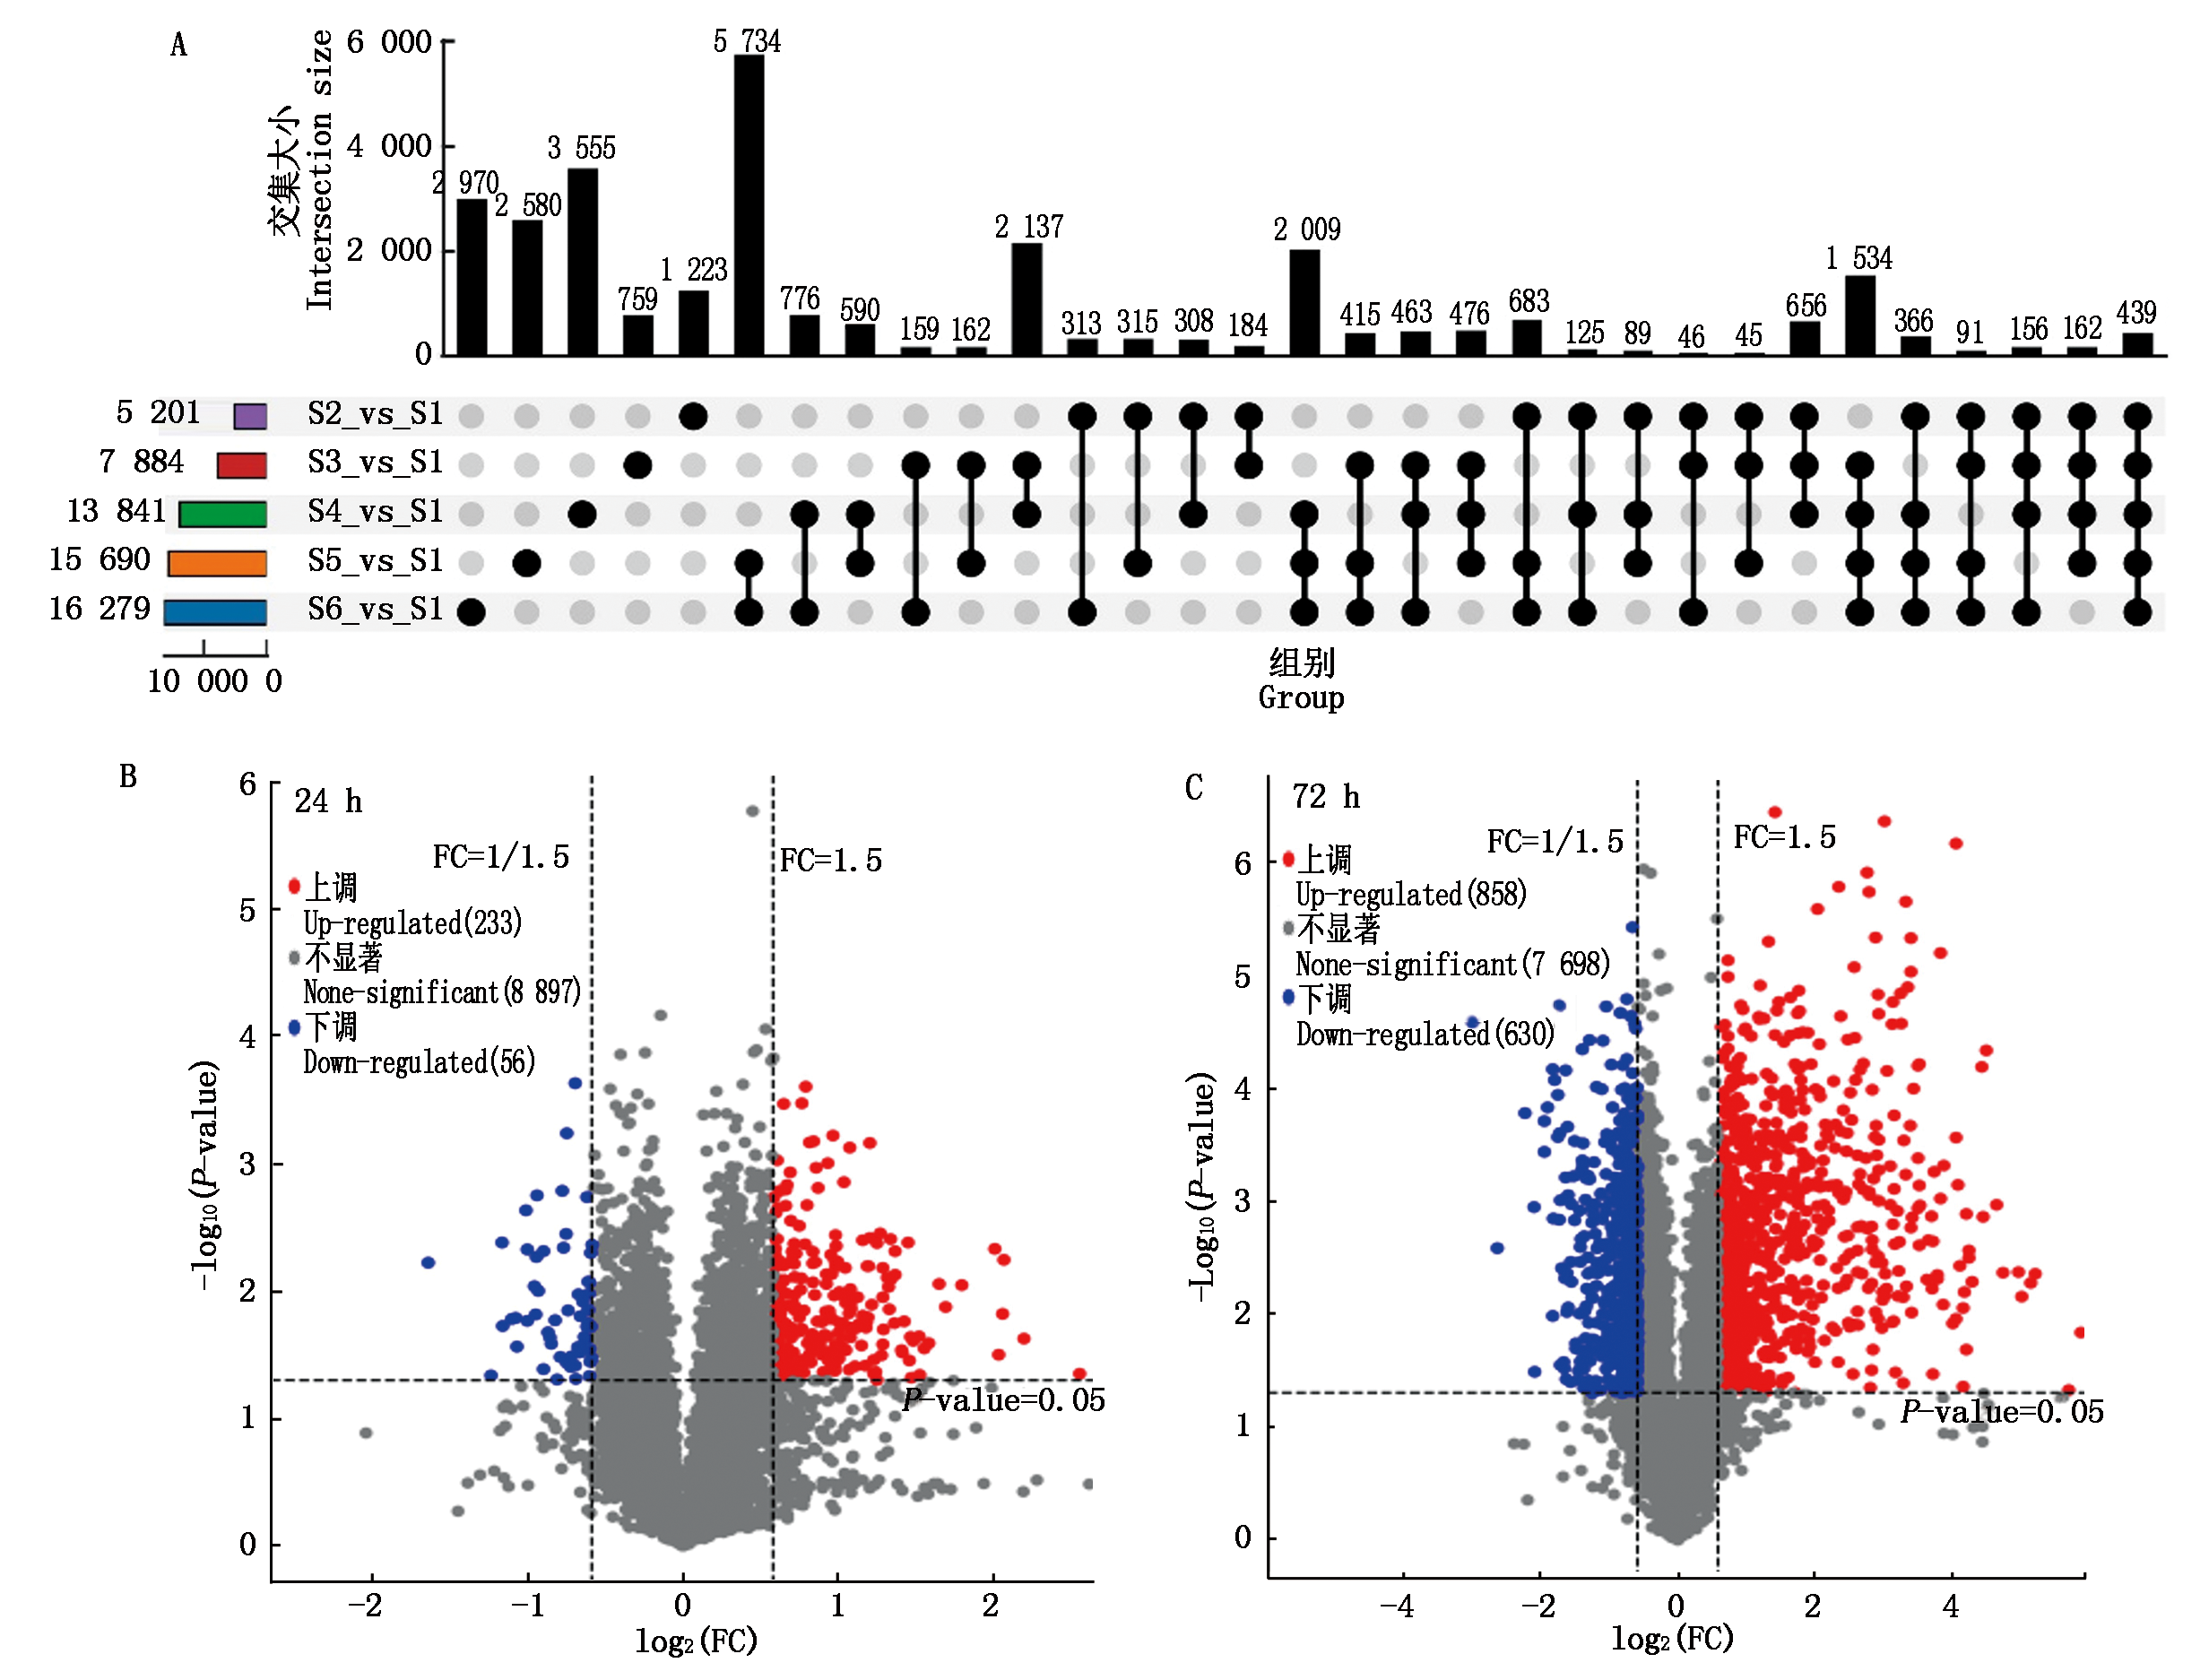
The identification of the wheat lamina joint regulator LIGULELESS 2 (LG2) and analysis of its interacting proteins are of great significance for exploring the molecular mechanisms underlying lamina joint formation and leaf angle regulation, as well as for breeding new varieties with ideal plant architecture. First, two homologous genes of LG2, designated TaLG2 and TaLG2L, were identified in the wheat genome through phylogenetic analysis, each containing three homoeologous copies located in the A, B, and D subgenomes with more than 90% sequence similarity. Transcriptome-based expression profiling during lamina joint development and in different leaf tissues revealed that both TaLG2 and TaLG2L were specifically expressed in the lamina joint. Except for TaLG2-3A, the other five homoeologs exhibited specific expression at the early S1 stage of lamina joint development. The bait vector pGBKT7-TaLG2L-1A was constructed and used for yeast two-hybrid (Y2H) screening yielding 174 positive clones.Sequencing analysis identified 68 candidate interacting proteins of TaLG2L-1A. GO enrichment analysis showed that these proteins were mainly involved in ubiquitin binding, cation binding, and mRNA binding. Further rotation verification and pairwise Y2H assays demonstrated that TaLG2L-1A can interact with the auxin response factor TaARF17 and the blue light receptor cryptochrome TaCRY1, respectively. This study clarifies the tissue-specific expression patterns of wheat LG2 homologs, identifies and obtains key interacting proteins TaARF17 and TaCRY1 of TaLG2L-1A, and reveals the molecular mechanism by which LG2 may coordinate auxin and light signaling pathways to regulate lamina joint formation and leaf angle development.

To investigate the key genes in cotton in response to Phytophthora boehmeriae infection and preliminarily elucidate the molecular mechanisms underlying their interaction,it performed transcriptome sequencing and TMT-based quantitative proteomic analysis on the boll rot-susceptible cotton germplasm line 885 using the Illumina HiseqTM platform.By screening for differentially expressed genes(proteins)and conducting GO/KEGG enrichment analyses,key genes involved in the stress response were identified.Bioinformatic analysis was further carried out on these candidate genes,and their expression patterns under pathogen stress were validated via Real-time Quantitative PCR(qPCR).The results showed that combined transcriptomic and proteomic screening identified 50 candidate sequences with significant differential abundance at both the transcriptional and translational levels.GO and KEGG enrichment analyses revealed that the differentially expressed genes were primarily enriched in pathways such as plant hormone signal transduction,plant-pathogen interaction,glutathione metabolism,and the biosynthesis of secondary metabolites.Further sequence alignment screened seven candidate genes related to disease resistance.qPCR verification demonstrated that the expression of Gh_D02G091500,Gh_D06G194600,Gh_A01G241100, Gh_A06G192400,and Gh_D09G161000 was significantly induced by P.boehmeriae.Among these,the cotton nitrate reductase gene exhibited the highest fold change and the most significant differential expression.The full-length sequence of this nitrate reductase gene was cloned and designated GhNIA1.In conclusion,it identified GhNIA1 as a key gene involved in the cotton-P.boehmeriae interaction and provided preliminary characterization of GhNIA1.These findings lay a theoretical foundation for further analysis of the molecular mechanism by which GhNIA1 participates in cotton resistance to boll rot and for breeding resistant varieties.
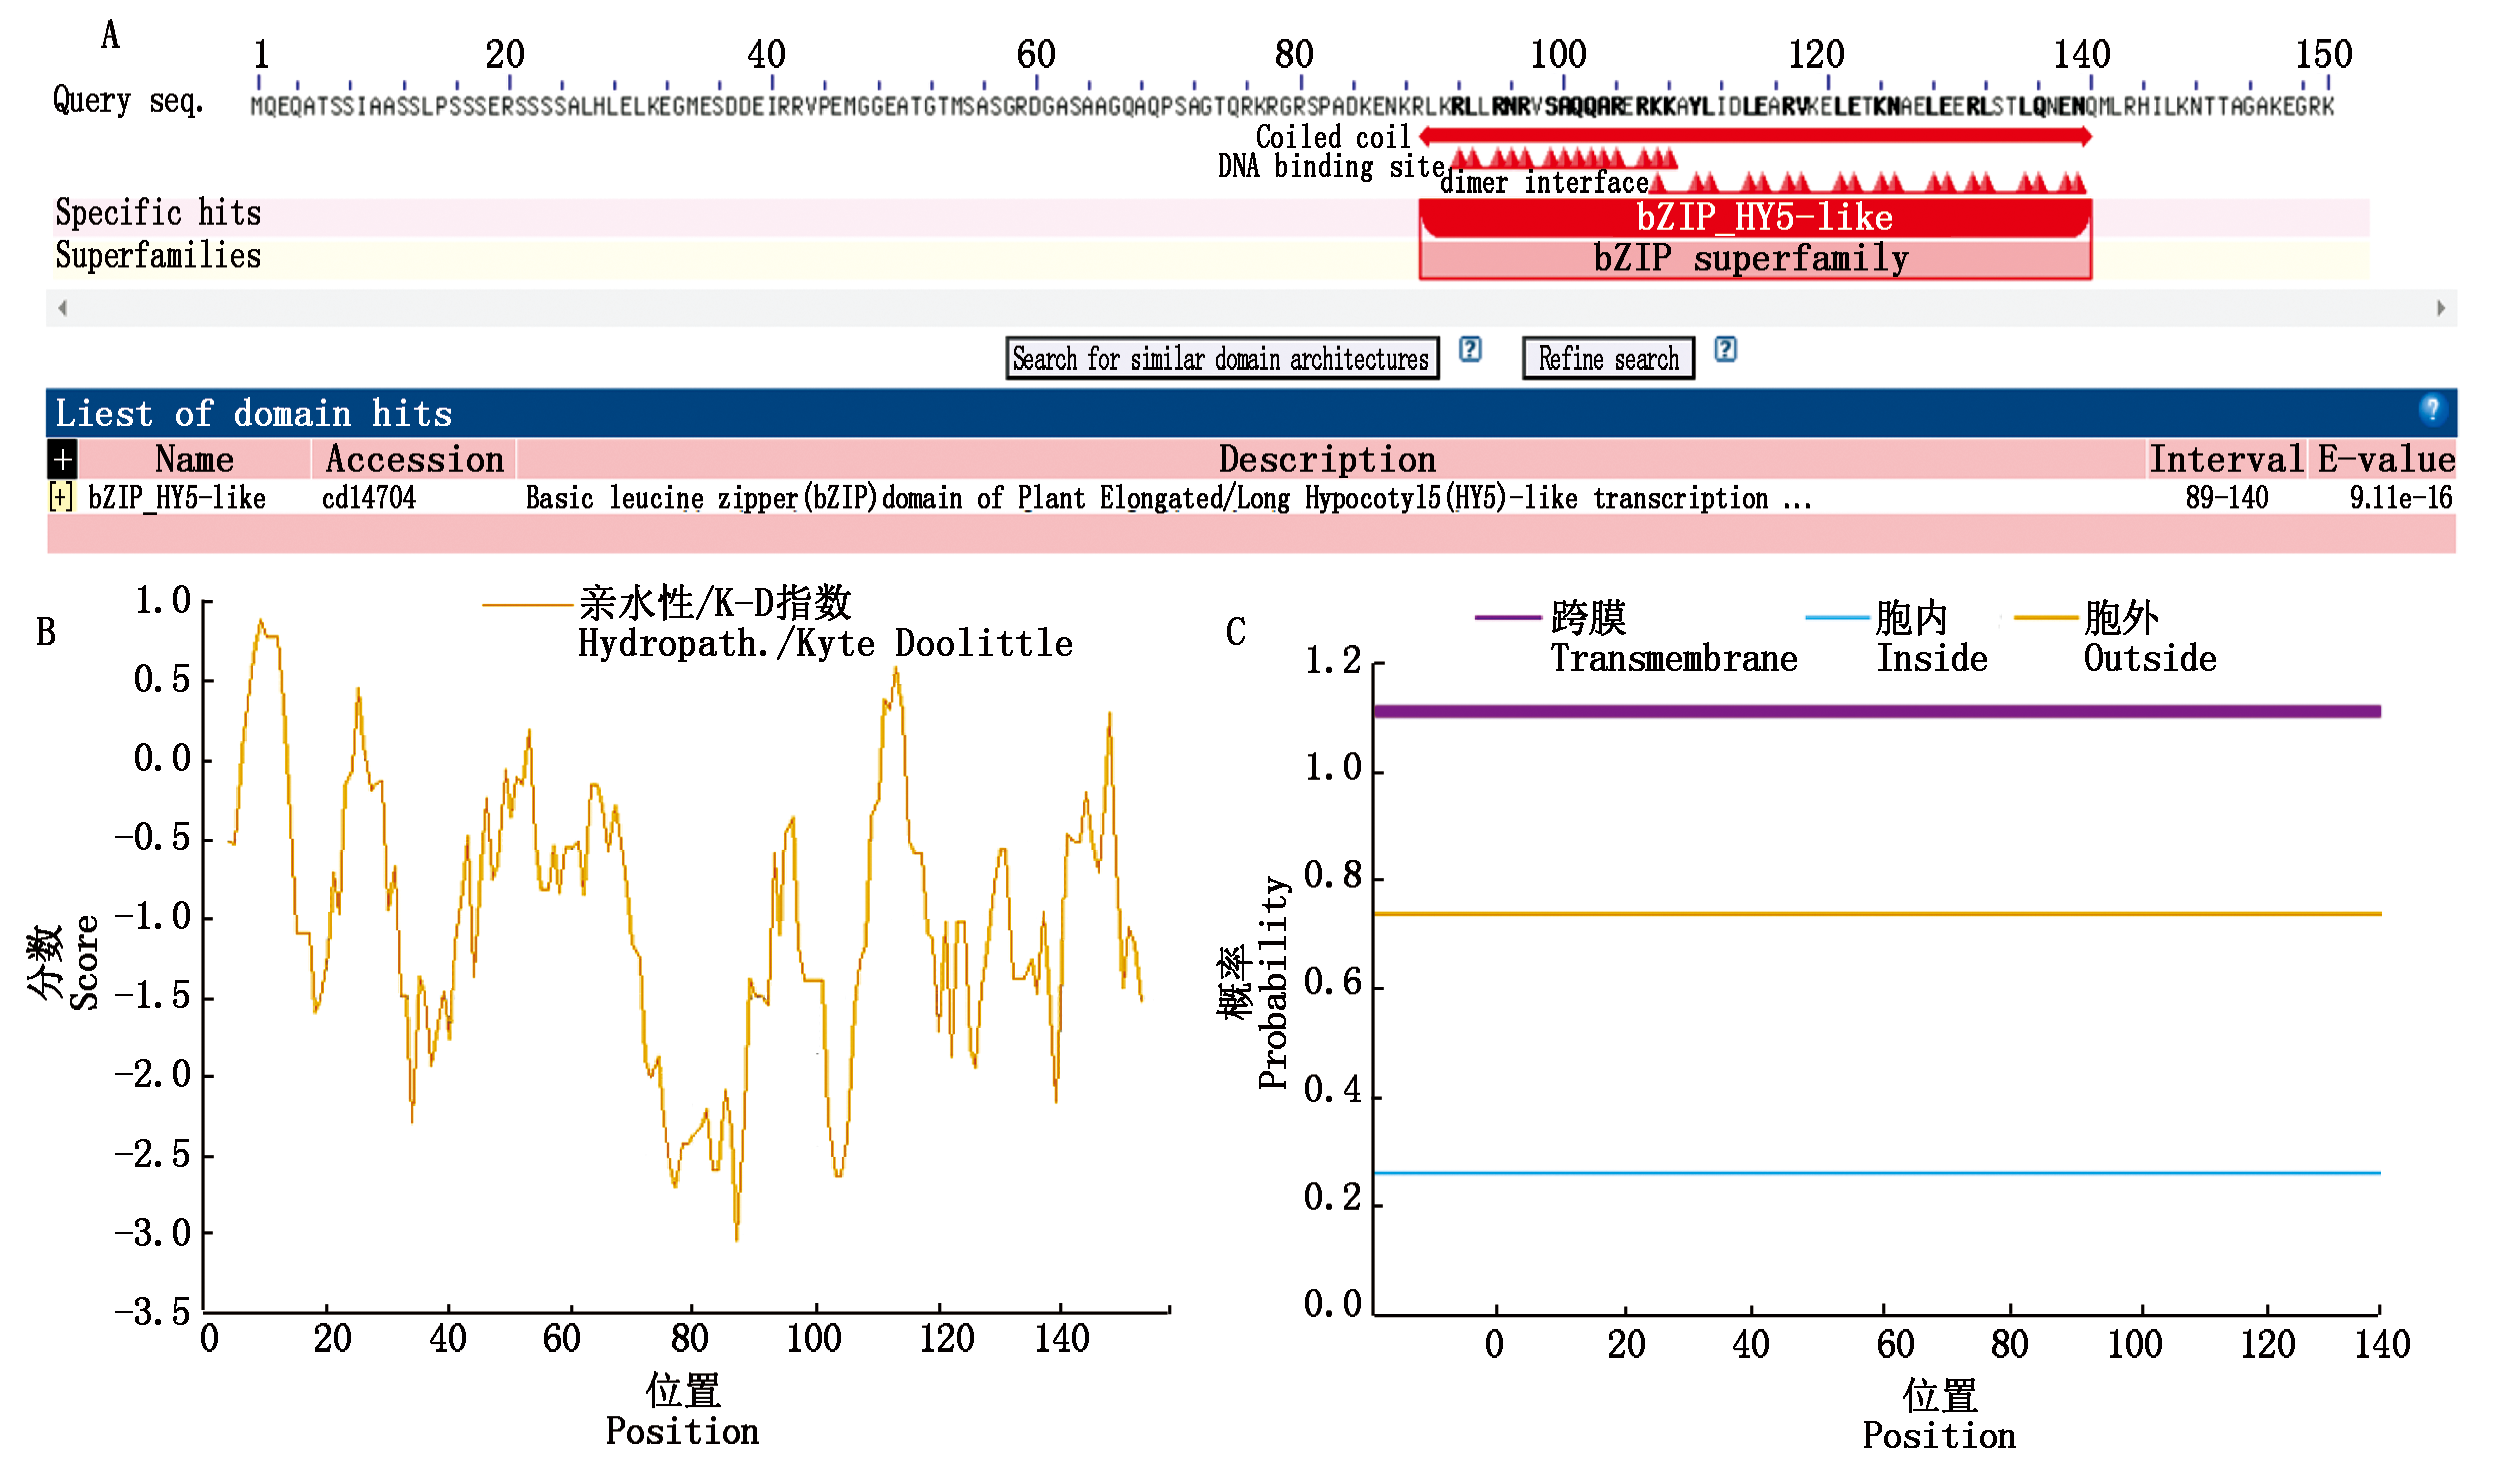
To clarify the basic characteristics and tissue expression pattern of the light signal transduction core bZIP transcription factor StHY5 gene,using potato cultivar Désirée as the experimental material,the full-length CDS sequence of the StHY5 gene was cloned.The sequence characteristics and evolutionary relationship were clarified by bioinformatics analysis.The expression patterns in different potato tissues were analyzed by Quantitative Real-time PCR(qRT-PCR).The results showed that the StHY5 protein consisted of 158 amino acids,with a molecular weight of 17.32 ku and a theoretical isoelectric point of 9.78.It contained one basic leucine zipper(bZIP)domain.The secondary structure of StHY5 was composed of 44.3% α-helices and 55.7% random coils,while the tertiary structure was dominated by α-helices,using the chicken CREB3 bZIP protein as the template.Phylogenetic tree analysis revealed that StHY5 clusters with HY5 proteins from Solanaceae species,showing the closest genetic relationship to CaHY5 in capsicum.qRT-PCR quantification demonstrated that StHY5 was expressed in all tested tissues with significant expression differences,among them the highest expression in stolons,and the lowest expression in tubers.This study clarifies the sequence characteristics and tissue expression pattern of the potato StHY5 gene,providing an experimental foundation for subsequent functional verification and pathway analysis in light signal regulation.
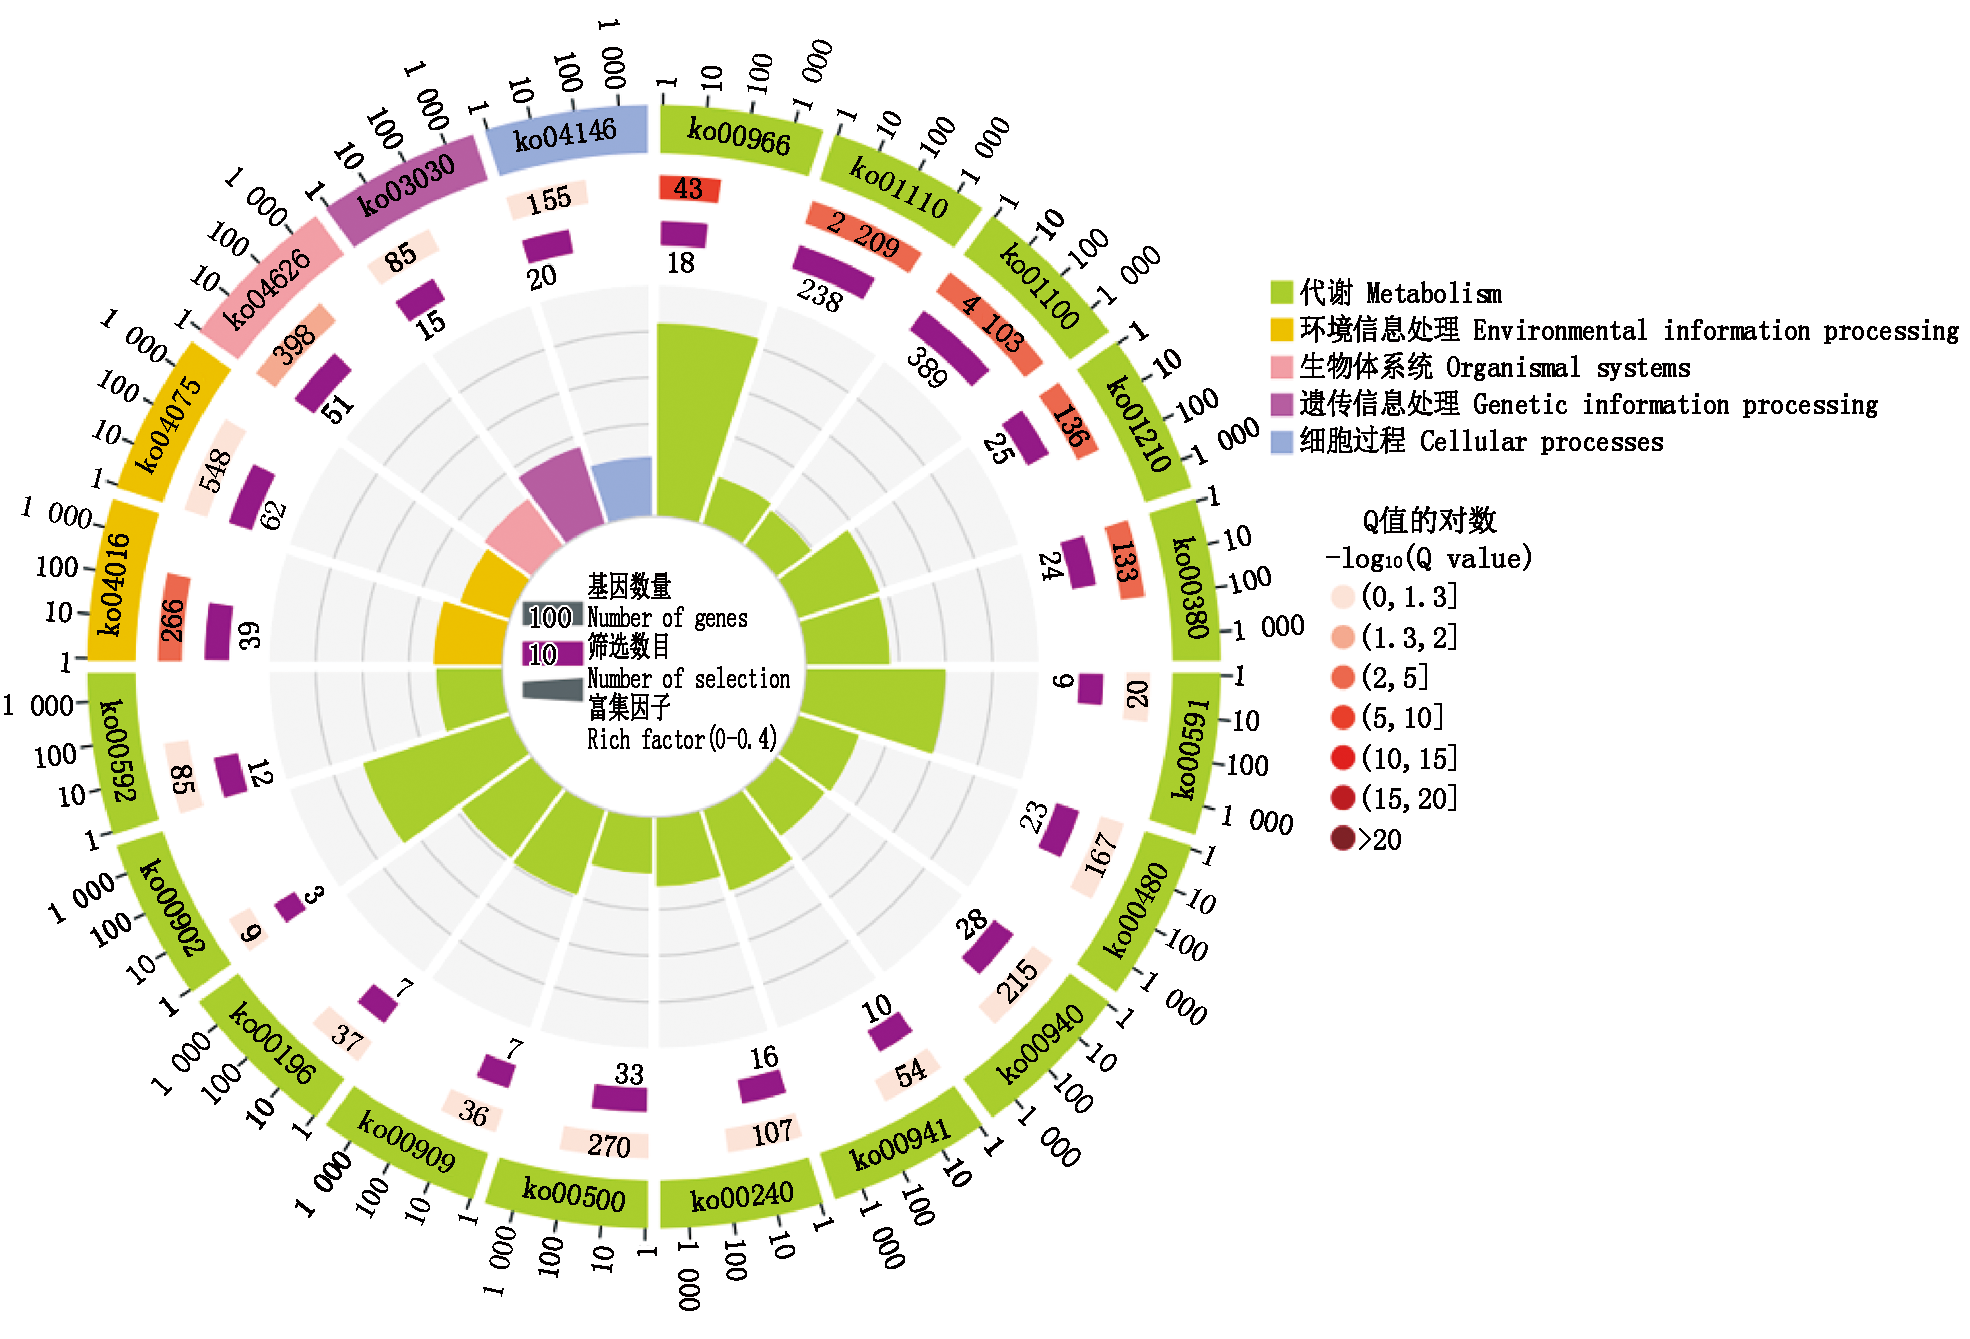
Flavonoids,as bioactive compounds in Chinese cabbage,possess antioxidant,anti-aging,and cancer-preventive properties.To investigate the molecular mechanisms underlying the differences in flavonoid accumulation among different ploidy Chinese cabbage materials,this study utilized 25 diploid and 25 tetraploid high-generation homozygous inbred lines.The total flavonoid content was systematically compared,and extreme lines were selected for transcriptome(RNA-Seq)analysis.Through KEGG pathway enrichment analysis,genes involved in flavonoid biosynthesis were identified,and RT-qPCR was employed to validate the expression patterns of differentially expressed genes and construct a transcription factor regulatory network.The results showed that the average total flavonoid content in tetraploid materials(10.96 mg/g)was slightly higher than that in diploid materials(10.36 mg/g).RNA-Seq analysis revealed 11 610 differentially expressed genes(DEGs)in diploid comparison groups and 6 659 DEGs in tetraploid comparison groups,with 3 598 common DEGs shared between them.KEGG enrichment analysis identified 34 candidate genes in the flavonoid and phenylpropanoid biosynthesis pathways.Six key genes(PAL1,4CL3,CHS3,CHI,CHI3,FLS1)were screened and found to be upregulated in high-flavonoid diploid and tetraploid materials.Among them,CHI3 expression was significantly higher in high-flavonoid tetraploid materials than in diploid materials.Additionally,27 bHLH and 19 MYB transcription factors were identified,and regulatory network analysis indicated that the MYB network was more complex and dominant than the bHLH network.RT-qPCR analysis confirmed that the expression patterns of the six selected key DEGs were consistent with the RNA-Seq data.This study identifies key structural genes and major transcription factors in the flavonoid biosynthesis pathway,revealing the important role of CHI3 in promoting flavonoid accumulation in tetraploid Chinese cabbage,thereby providing candidate genes and a theoretical foundation for breeding high-flavonoid Chinese cabbage.
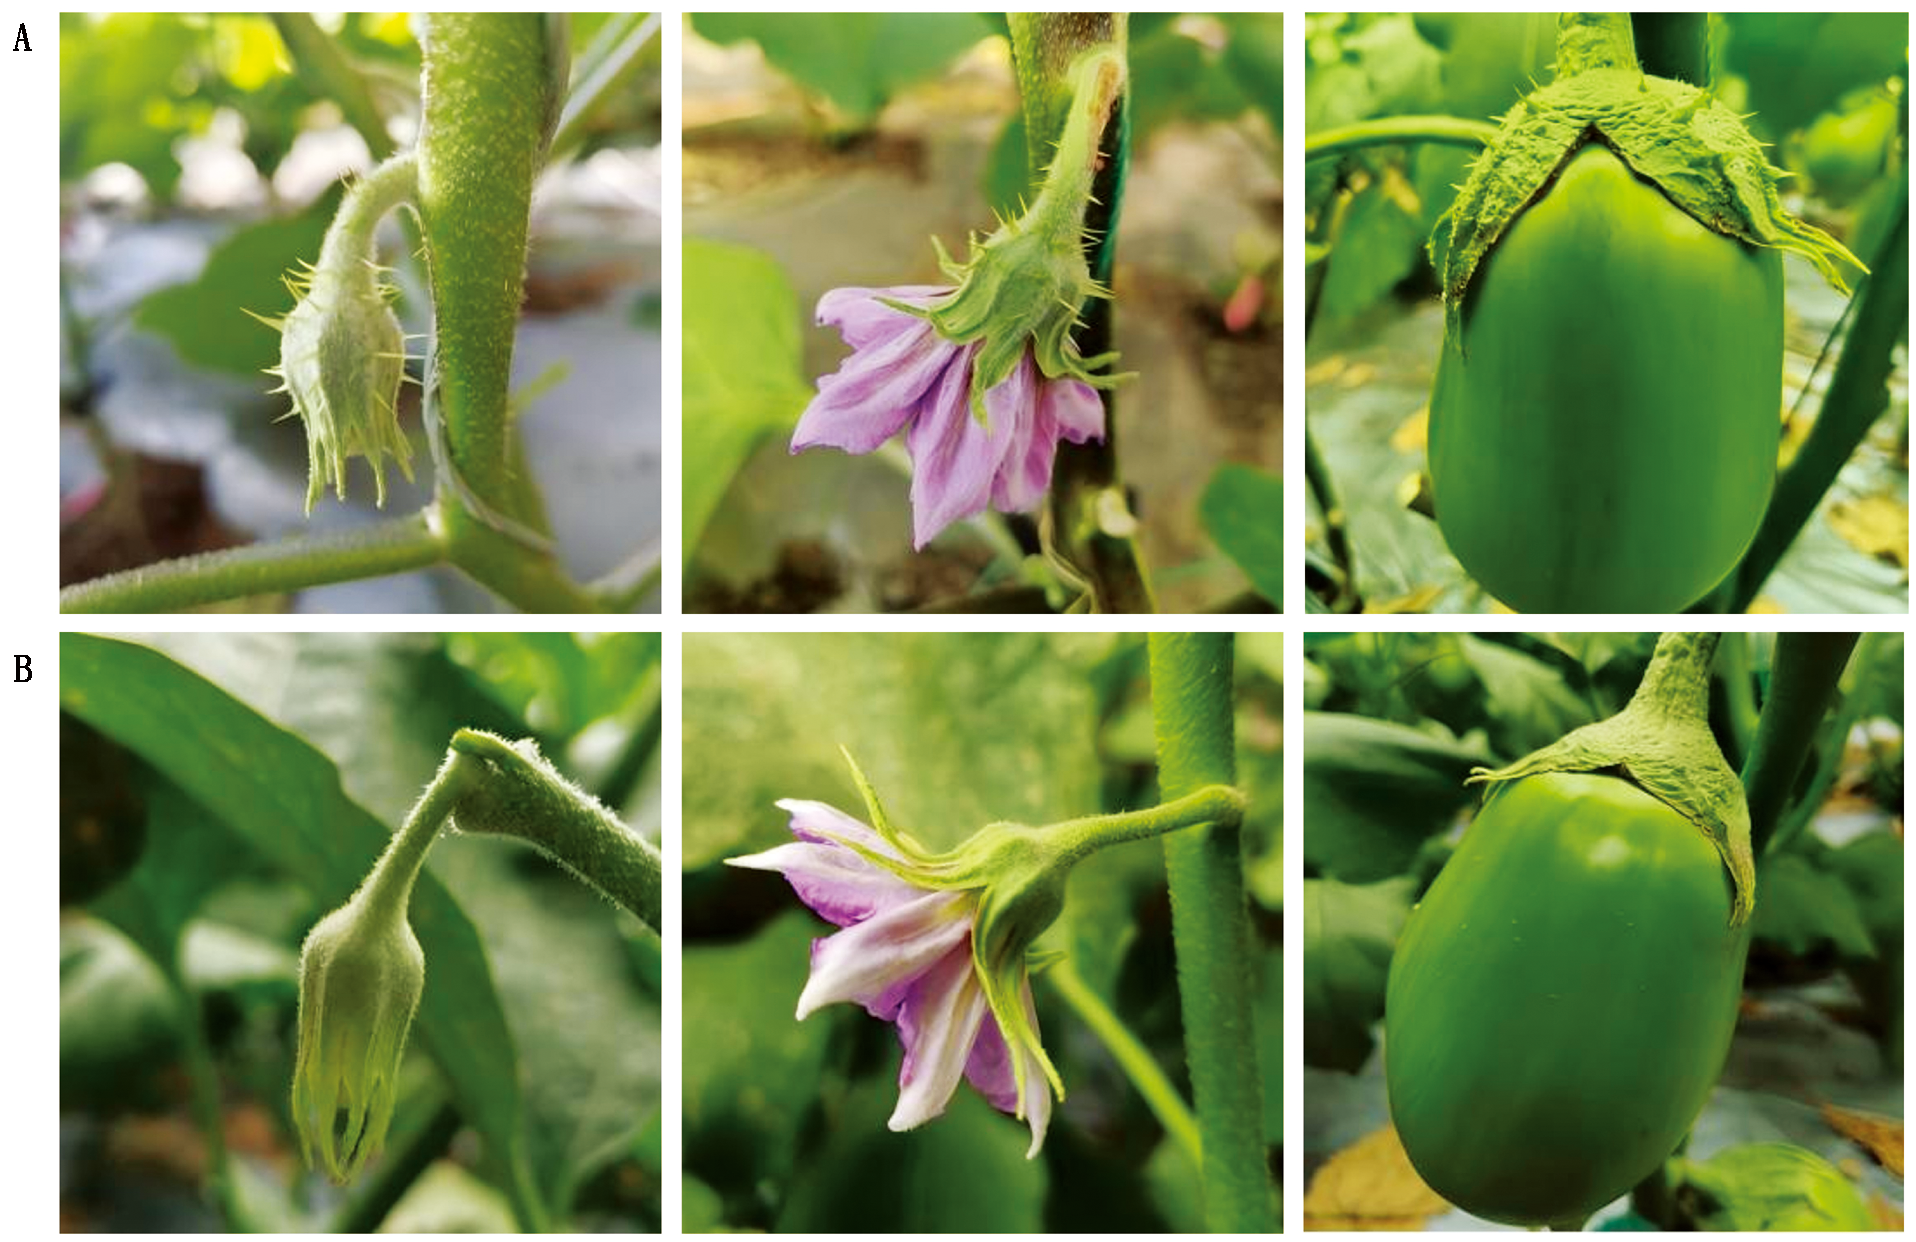
To elucidate the molecular mechanism of prickle formation on the eggplant calyx,a genetic population was developed by crossing the homozygous prickly line 24C01(female parent)and the non-prickly line 24C02(male parent).In the F2 generation,the distribution of calyx prickle density followed a normal distribution.Two extreme bulks were constructed by pooling DNA from 30 highly prickly and 30 completely non-prickly F2 individuals.By combining BSA-seq with map-based cloning,the calyx prickle gene SmPL was fine-mapped to a 131.4 kb interval on chromosome 6.Among the 12 annotated genes within this region,only Smechr0602815.1,Smechr0602817.1,and Smechr0602821.1 showed differential expression between the two parental lines.Re-sequencing analysis further revealed a nonsynonymous mutation(G1066A)in Smechr0602815.1 in 24C02,resulting in a glutamate-to-lysine substitution,and a fragment deletion in Smechr0602821.1 in the same parent.Based on these sequence variations,kompetitive allele-specific PCR(KASP)markers linked to the non-prickly allele were developed.It was found that the markers exhibited a genotype-phenotype concordance rate exceeding 96% when tested across 100 diverse eggplant accessions.These results indicate that the developed KASP markers enable efficient early-selection for the non-prickly trait and can significantly accelerate the breeding of nonprickly eggplant varieties.
To explore the formation molecular mechanism of the geoherbalism of Shanxi Forsythia suspensa,a genome-wide identification and characterization of UGT gene family members was performed using bioinformatics methods based on the genome data of F.suspensa.Transcriptome data of F.suspensa from different producing areas and Real-time PCR were used to analyze the expression patterns of UGTs gene in F.suspensa from different habitats and different tissues.The results showed that a total of 170 family members were identified in the whole genome of F.suspensa,and the family members were unevenly distributed on 14 chromosomes,named as FsUGT1—FsUGT170.The molecular weight of 11.16—56.77 ku and isoelectric point of 4.70—9.18,most of which are hydrophilic proteins.Subcellular localization predictions indicated that FsUGT proteins were predominantly localized in the chloroplast and cytoplasm.Phylogenetic analysis classified the F.suspensa UGT gene family into 15 distinct subgroups.Synteny analysis identified 31 collinear gene pairs involving 23 FsUGTs and 20 UGTs from Arabidopsis thaliana.Analysis of gene structure,conserved protein motifs,and conserved domains revealed that the FsUGT genes possess 1 to 7 exons and 0 to 6 introns,and all encoded proteins contain the characteristic PSPG conserved domain.Promoter cis-acting element analysis demonstrated the presence of multiple regulatory elements associated with light responsiveness,phytohormone signaling,biotic and abiotic stress responses,and growth and development within the promoter regions of FsUGT genes.Gene expression profiling showed that the transcript levels of FsUGT4 and FsUGT145 were significantly higher in F.suspensa samples from Shanxi compared to those from Henan and Shaanxi,and their expression in leaves was significantly higher than in flowers and fruits.In conclusion,this study identified 170 members of the UGT gene family in F.suspensa and highlighted two genes,FsUGT4 and FsUGT145,which exhibit elevated expression in Shanxi accessions and are potentially implicated in forsythin biosynthesis.

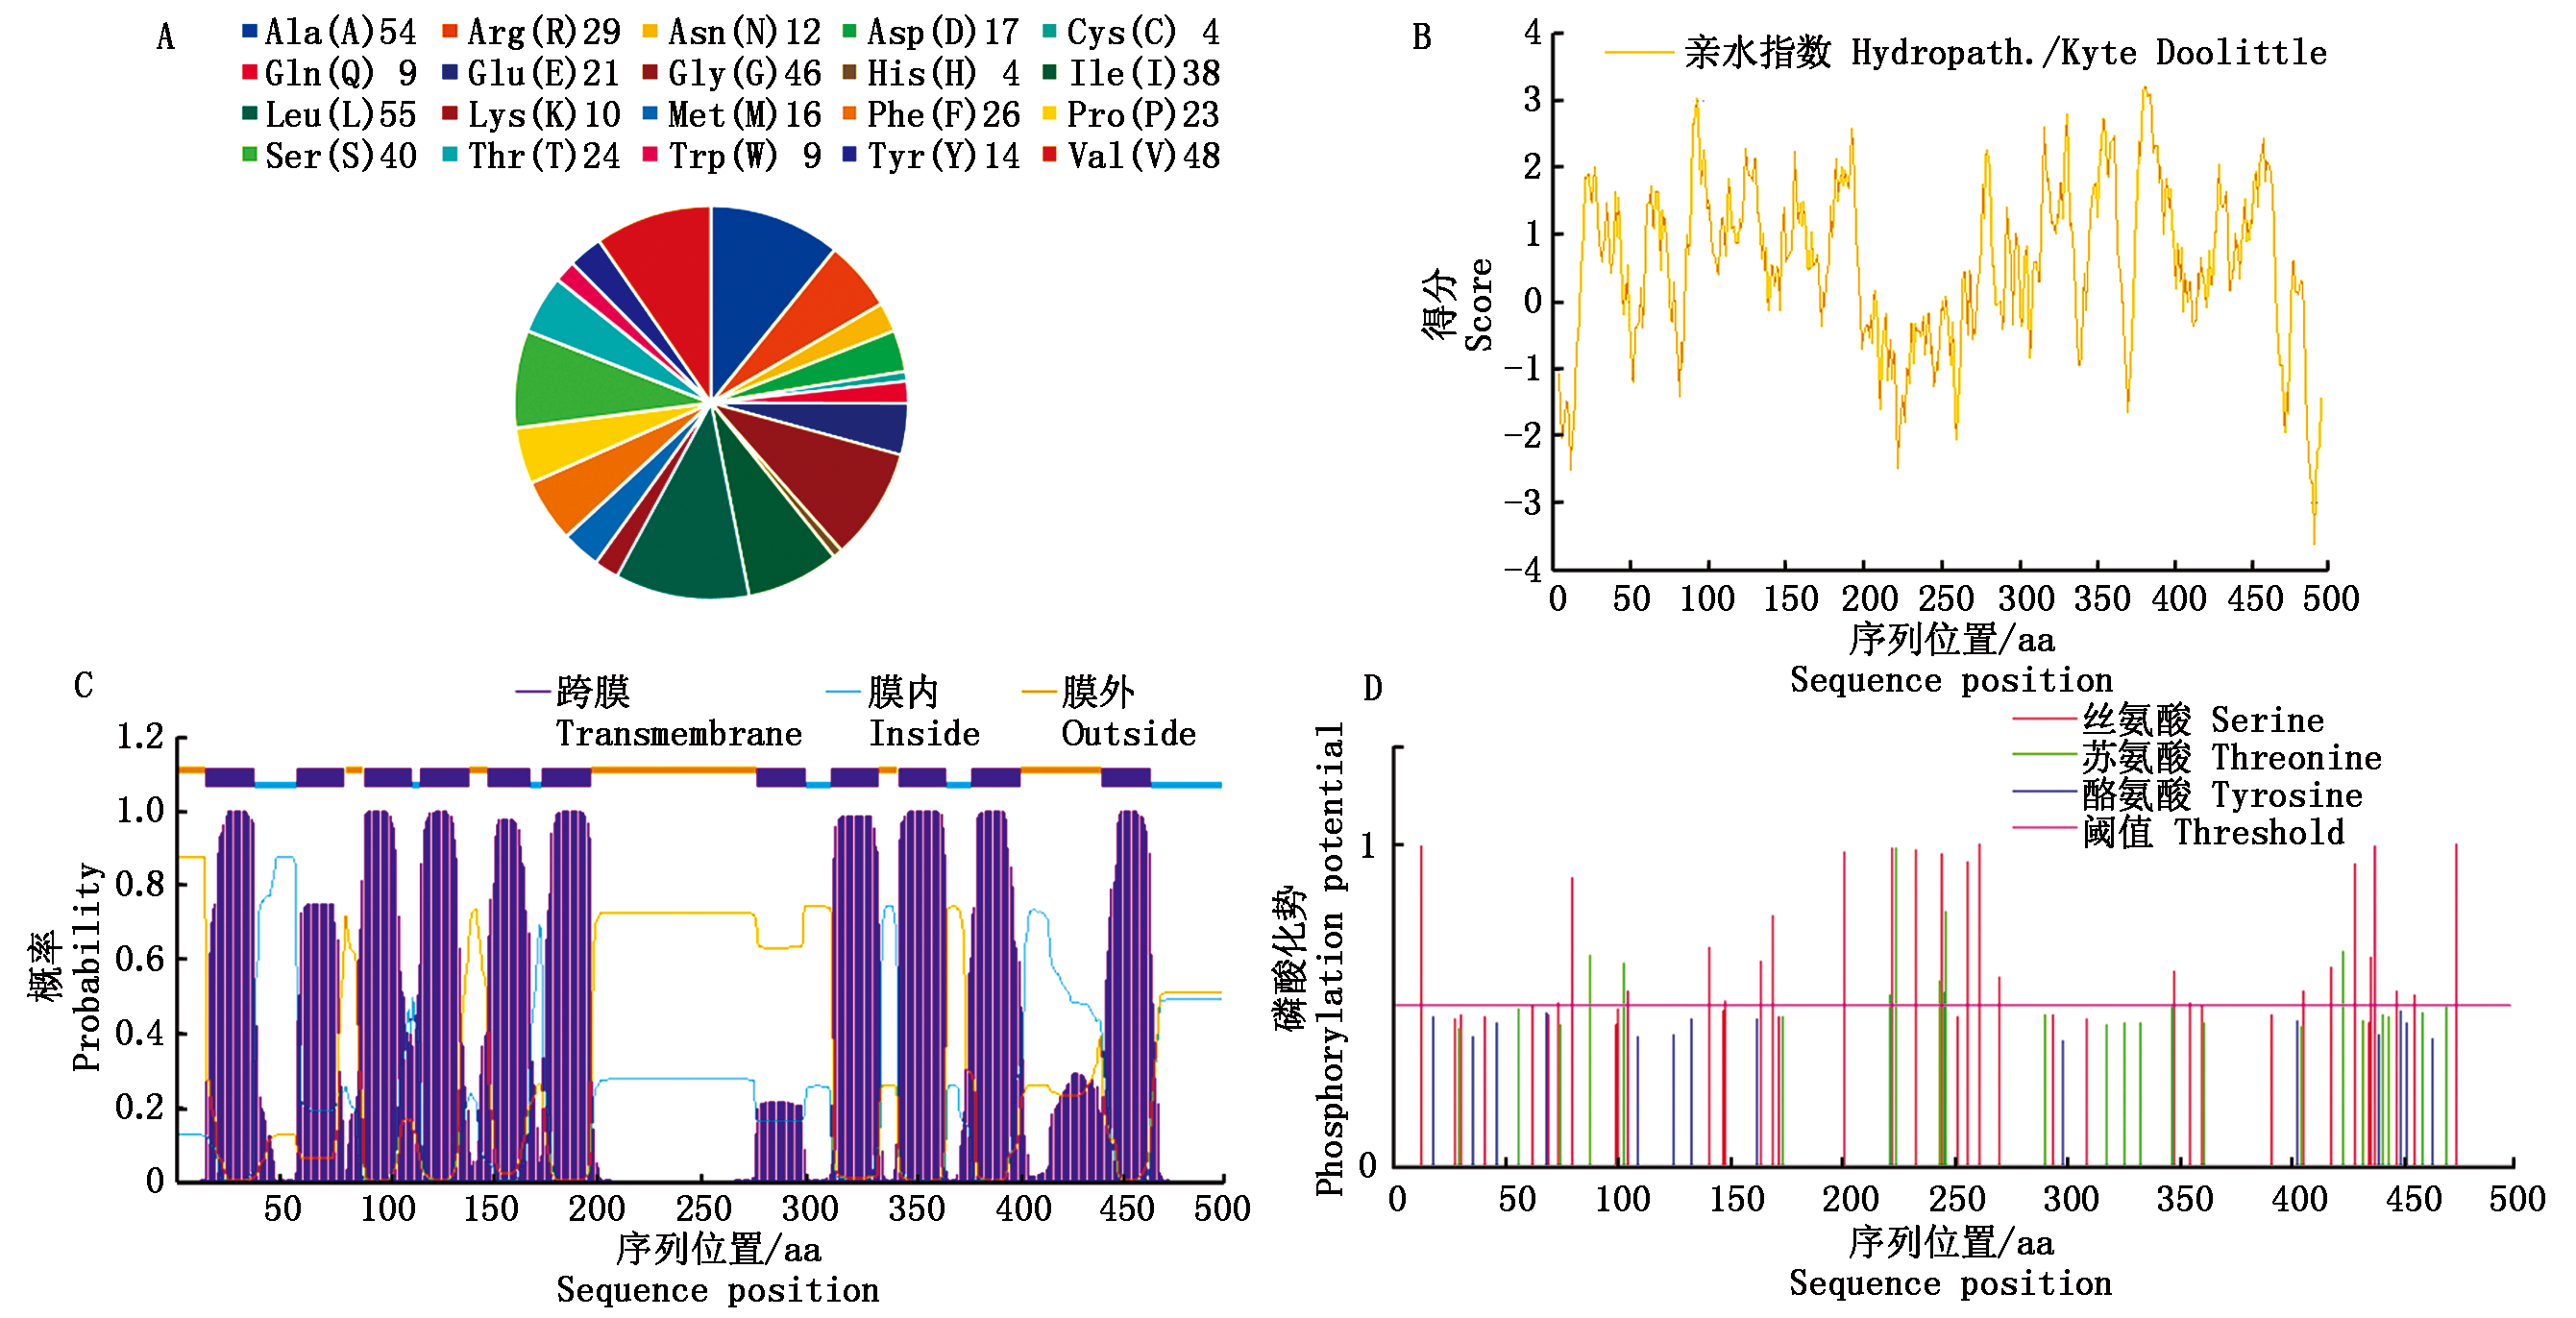
Bulblet enlargement is of significant importance for lily cultivation and breeding.The PLETHORA(PLT)transcription factors play a key regulatory role in plant organ expansion.This study used Lilium Tresor as experimental material to investigate the mechanism of the LaPLT5 gene during its bulblet enlargement process.By analyzing transcriptome data from small bulblets at stages S1 to S5 of Lilium Tresor,the LaPLT5 gene,which is potentially involved in bulblet enlargement,was identified.The gene was cloned from bulblet cDNA using molecular cloning techniques,and the structural characteristics of its encoded protein were predicted through bioinformatics analysis.The expression pattern of LaPLT5 during bulblet enlargement was examined using qRT-PCR,and its function was validated via virus-induced gene silencing(VIGS)and transient overexpression assays.The results showed that the LaPLT5 gene had a full length of 1 500 bp,encoding 499 amino acids,including 39 positively charged residues and 38 negatively charged residues.The protein had a relative molecular weight of 53 754.25,an instability index of 40.77,and an average hydrophilicity value of 0.56,indicating that it is an unstable hydrophobic protein.qRT-PCR analysis revealed that LaPLT5 expression was highest in the middle scale layers of Lilium Tresor bulblets,followed by that in small bulblets,and its expression level continuously increased from stages S2 to S4 during the enlargement of small bulblets.After VIGS-mediated silencing of LaPLT5,the diameter,circumference,and weight of small bulblets were highly significantly greater than those in the control group,whereas transient overexpression of LaPLT5 yielded opposite results.This indicated that LaPLT5 negatively regulated the enlargement of small bulblets in Lilium Tresor.Based on the phenotype of increased cell number but reduced cell area in small bulblets after LaPLT5 silencing,qRT-PCR analysis showed that the expression of cell wall relaxation-related genes,LaPME34,LaXTH1 and LaXTH6,was highly significantly downregulated.Meanwhile,the positive cell wall regulatory genes LaEBP1 and LaANT1 were highly significantly upregulated, whereas the negative regulatory gene LaCKX4 was highly significantly downregulated.These findings suggested that LaPLT5 likely regulated the expansion and division of small bulblet cells by coordinating the expression of genes involved in cell wall remodeling and cell division.In summary,LaPLT5 participates in the enlargement of small bulblets in Lilium Tresor and functions as a negative regulator of this process.
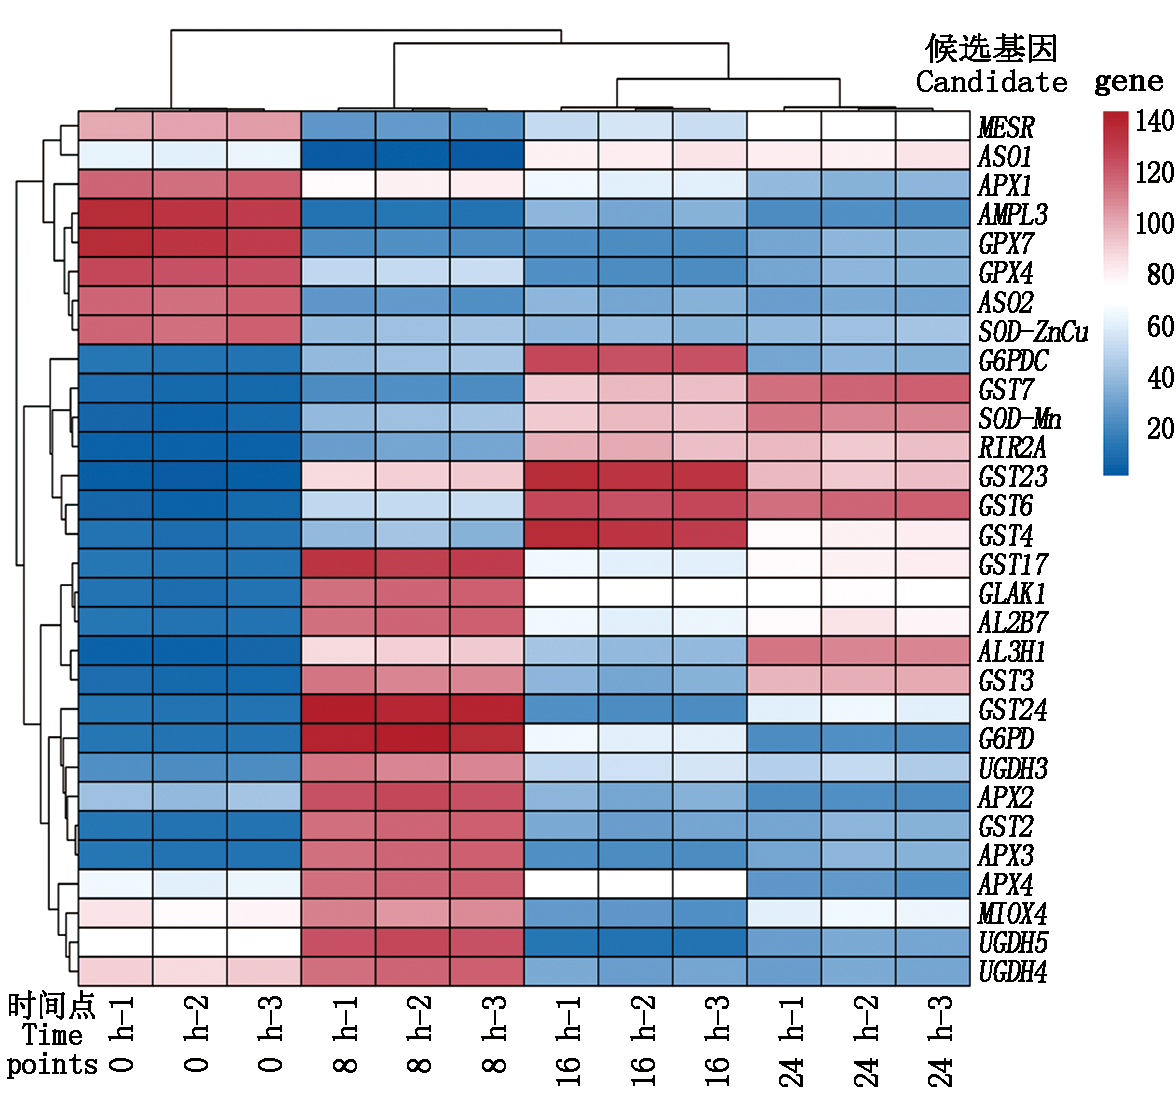
In order to uncover key genes involved in flax's response to drought stress and elucidate the underlying molecular mechanisms,thereby providing theoretical support for breeding drought-resistant varieties.Using roots of the drought-resistant flax cultivar Tianya 13 as experimental material,drought stress was simulated with different concentrations(5%,10%,20%,30%)of polyethylene glycol(PEG).Based on the analysis of physiological indicators such as root length,root thickness,and dry/fresh weight ratio,20% PEG was identified as the optimal stress concentration,under which the above indicators were significantly reduced by 31.63%,30.50%,and 13.39 percentage points,respectively,compared to the control.Subsequently,Illumina transcriptome sequencing was performed on roots subjected to drought stress at different time points(8,16,24 h),identifying 5 428,5 304,and 3 603 differentially expressed genes(DEGs)at each time point,respectively.KEGG enrichment analysis revealed that these DEGs were significantly enriched in key pathways such as carbohydrate metabolism(12 genes),betaine biosynthesis(4 genes),and the antioxidant defense system(36 genes).qRT-PCR validation of selected DEGs confirmed expression trends consistent with the RNA-seq results,verifying data reliability.This study systematically reveals,that flax enhances drought tolerance through the coordinated regulation of osmotic adjustment substance(e.g.,betaine)synthesis,energy metabolism reprogramming(carbohydrate metabolism),and reactive oxygen species scavenging mechanisms.It provides important theoretical foundations and candidate gene resources for molecular design breeding in flax.
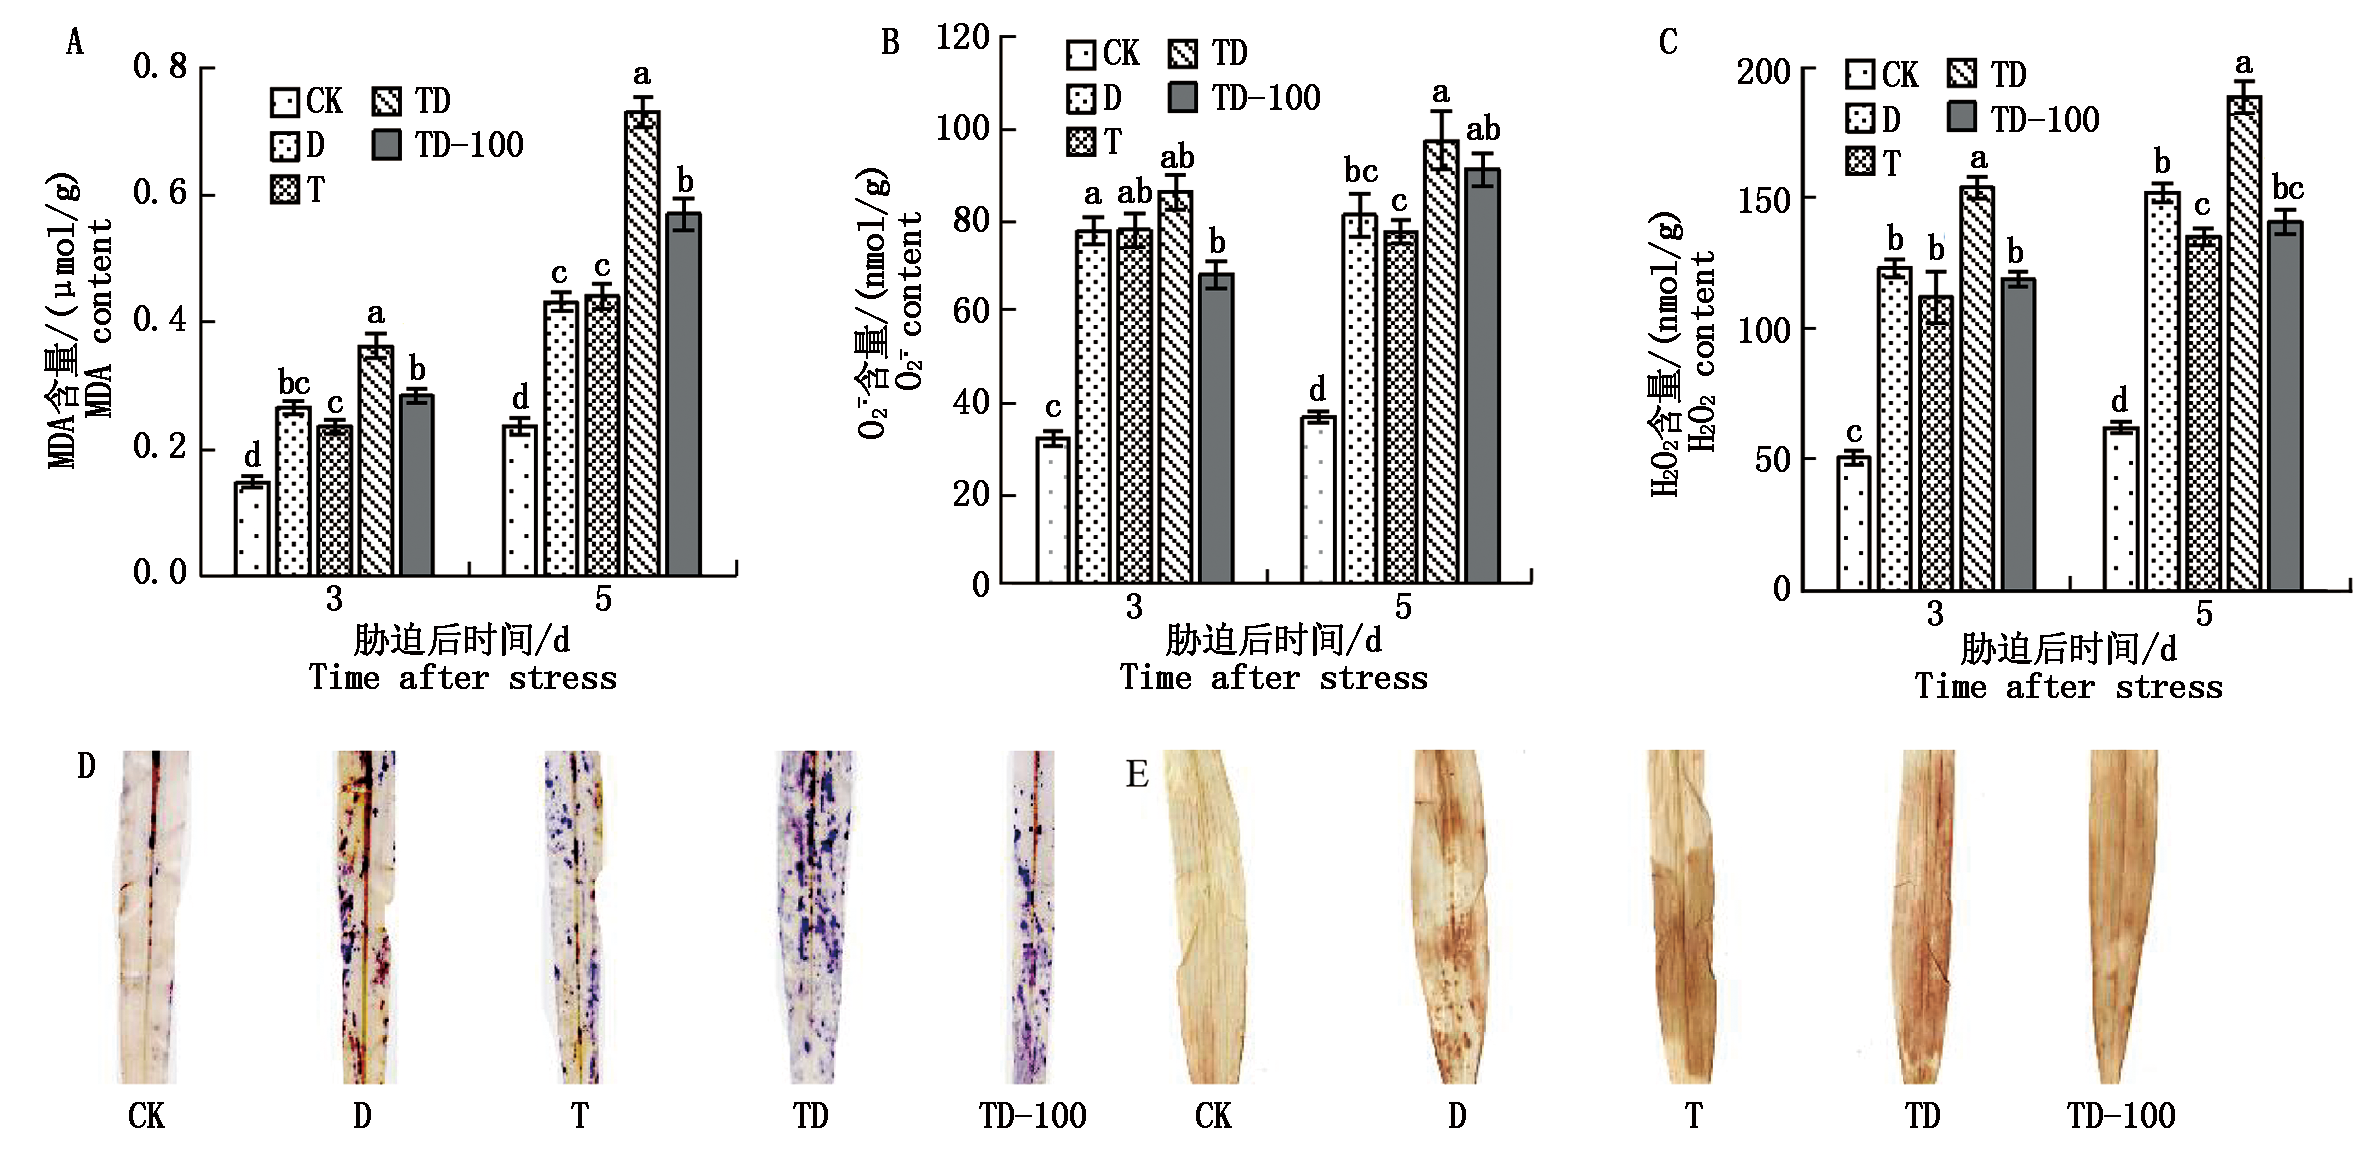
This study aimed to clarify the regulatory mechanism of exogenous glycine betaine(GB)on the antioxidant and photosynthetic systems of maize seedling leaves under combined heat and drought stress(TD),providing a theoretical basis for alleviating crop stress under extreme environments.Using the maize variety Xianyu 335 as experimental material,different concentrations of GB(0,50,100,200,and 400 mmol/L)were applied to the leaves at the three-leaf-one-heart stage.After 24 hours,combined stress was simulated in a climate chamber with high temperature(day/night temperature of 40 ℃/28 ℃)and 20% PEG-6000 to induce moderate drought,exploring the regulatory effects of GB on the antioxidant system and photosynthesis of maize seedlings.The results showed that 100 mmol/L GB significantly promoted plant growth.Compared with the TD treatment,plant height,leaf area,shoot dry weight,and root dry weight increased by 27.48%,27.45%,28.33%,and 78.26%,respectively,while the root-to-shoot ratio increased by 14.91 percentage points.On the 5th day after stress,100 mmol/L GB significantly enhanced antioxidant enzyme activities,with superoxide dismutase(SOD),peroxidase(POD),and catalase(CAT)activities increasing by 69.09%,69.02%,and 41.81%,respectively.Simultaneously,the contents of malondialdehyde(MDA),superoxide anion($\mathrm{O}^{\bar{.}}_{2}$),and hydrogen peroxide(H2O2)decreased by 21.92%,6.50%,and 25.33%,respectively.Furthermore,soluble sugar(SS)and soluble protein(SP)contents increased by 31.69% and 18.98%.Photosynthetic characterization analysis indicated that GB effectively improved the lamellar system structure of chloroplast grana and stroma thylakoids,thereby significantly increasing the SPAD value under combined stress and alleviating damage to the photosystem Ⅱ(PSⅡ)reaction center.This was manifested by a decrease in minimal fluorescence(F0)and increases in variable fluorescence(Fv)and maximum fluorescence(Fm).Additionally,by mitigating the stomatal and non-stomatal limitations caused by combined stress,GB treatment significantly enhanced photosynthetic performance,increasing the net photosynthetic rate(Pn)by 24.14% on the 5th day after stress.In conclusion,under combined heat and drought stress,exogenous GB can protect the photosynthetic system by enhancing antioxidant enzyme activities and osmotic adjustment capacity,thereby alleviating stress damage and promoting the growth of maize seedlings.
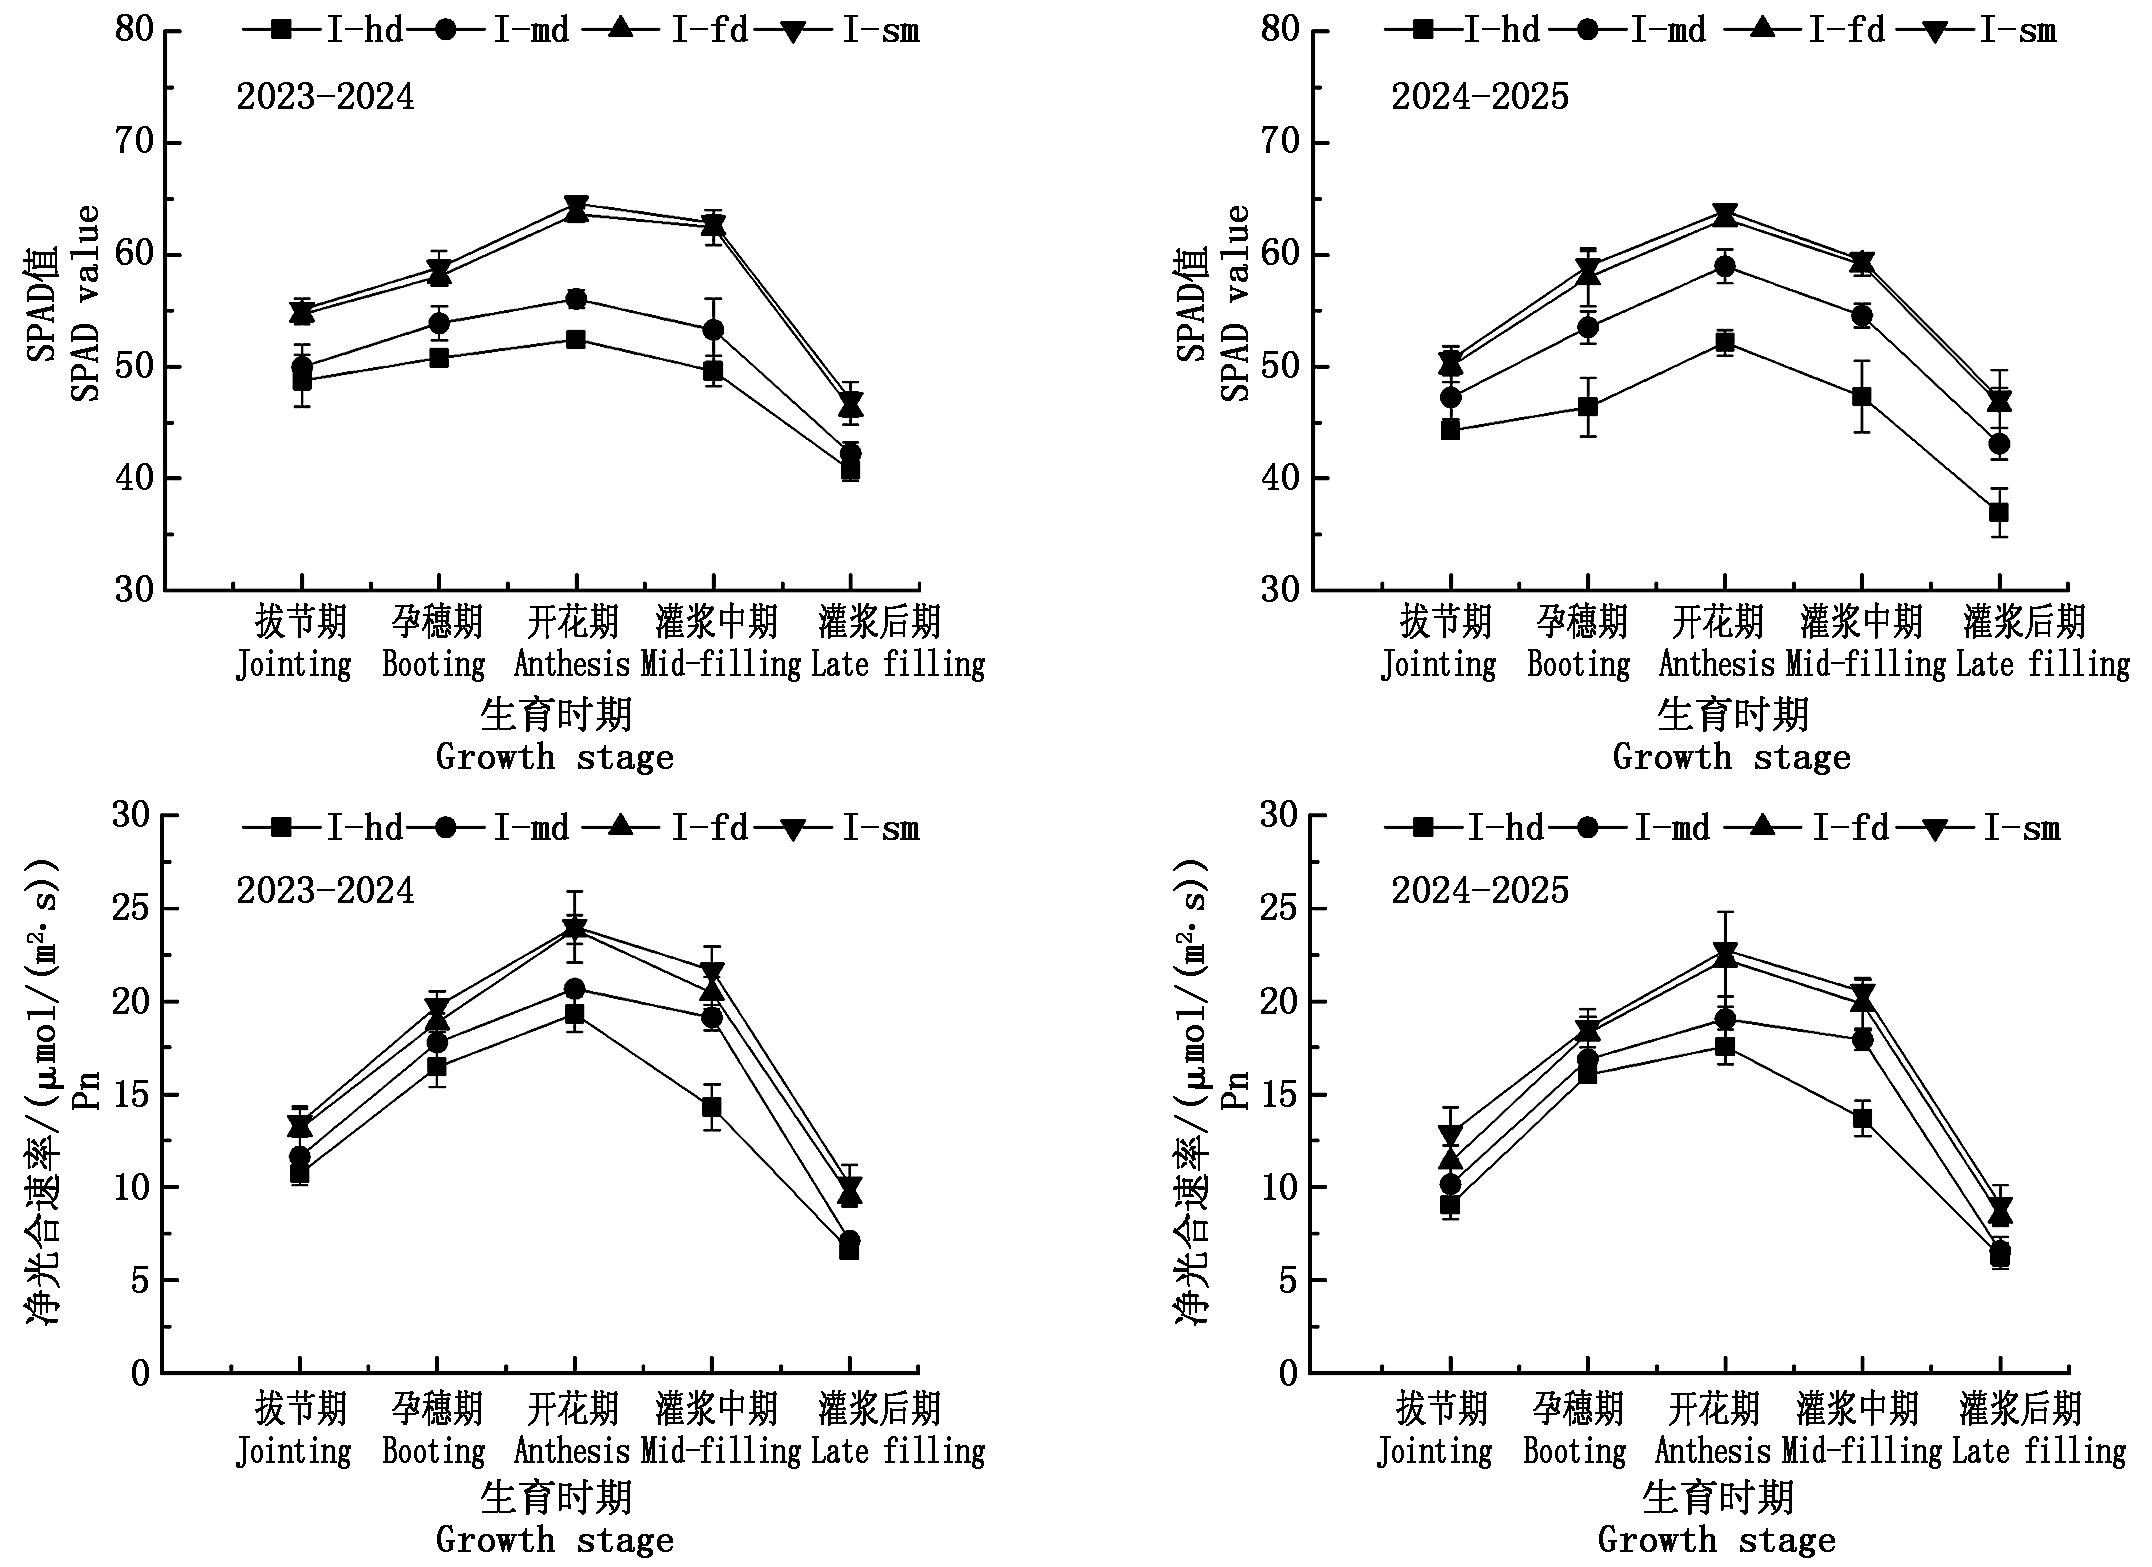
The effects of different degrees of drought stress on antioxidant enzyme activity, malondialdehyde(MDA)content,photosynthetic characteristics,and yield composition of wheat were investigated to provide a theoretical basis and technical support for optimizing water-saving production of wheat.From 2023 to 2025,four soil water gradients of heavy drought(I-hd),moderate drought(I-md),mild drought(I-fd)and suitable water content(I-sm)were set up in the Longkou Experimental Station of Shandong Academy of Agricultural Sciences.The antioxidant enzyme activity,MDA content,photosynthetic characteristics and grain yield composition of wheat under different drought stress were systematically analyzed.The results showed that compared with moderate and heavy drought,mild drought and suitable water were beneficial to optimize spike number,grain number per spike and 1000-grain weight,thus significantly increasing grain yield,and the average grain yield in two years increased by 11.1%—51.0%.The activities of antioxidant enzymes such as superoxide dismutase(SOD),peroxidase(POD),and catalase(CAT)were significantly increased at each growth stage under mild drought and suitable water,and the MDA content in leaves was significantly decreased at each growth stage,which played an important role in reducing the degree of cell membrane lipid oxidation and delaying plant senescence.The leaf area index(LAI),chlorophyll relative value(SPAD value)and net photosynthetic rate(Pn)of leaves at booting,flowering and mid-filling stages were maintained at a high level under mild drought and suitable water conditions,which provided a physiological basis for the improvement of photosynthetic rate,while the above indices were significantly reduced during the whole growth period under moderate and severe drought conditions.The results of correlation analysis showed that the activities of antioxidant enzymes(SOD,POD,CAT)were significantly negatively correlated with MDA content,and significantly positively correlated with photosynthetic characteristics(LAI,SPAD,Pn)and yield components(spike number,grain number per spike,1000-grain weight),which provided physiological guarantee for maintaining higher photosynthetic capacity.In summary,both mild drought and suitable water significantly increase the activities of antioxidant enzymes such as SOD,POD,and CAT in leaves at each growth stage,reduce the content of harmful substances such as MDA,and optimize LAI and SPAD values to improve the photosynthetic rate of leaves.The final grain yield increased by 11.1%—51.0%.However,there was no significant difference in grain yield,antioxidant enzyme activity and photosynthetic characteristics between mild drought and suitable water.Therefore,it is concluded that under mild drought conditions,high and stable yield of wheat can be achieved by optimizing photosynthetic characteristics and leaf antioxidant capacity,which provides a reliable practical approach for water-saving crop production.
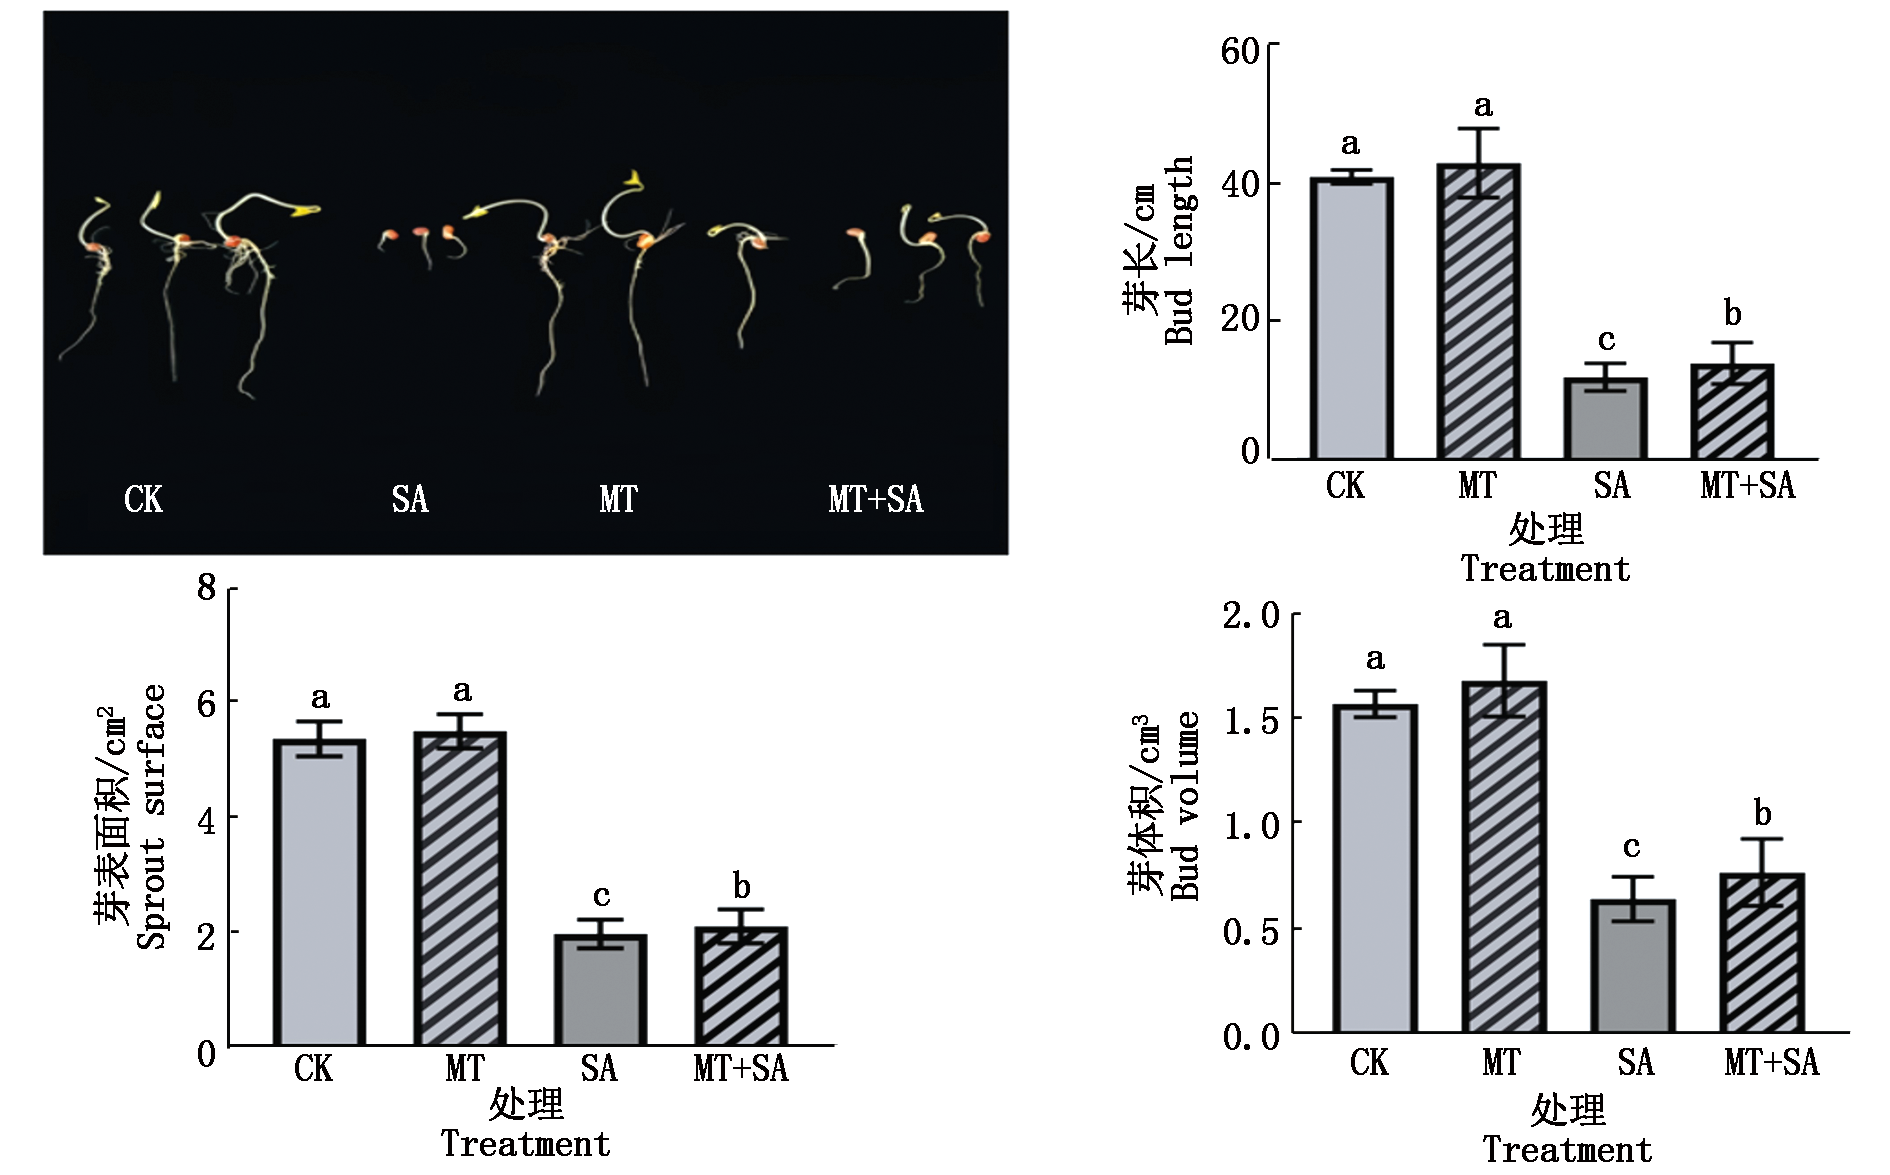
Excessive salt-alkali in the soil will inhibit the growth of adzuki bean plants,resulting in a decrease in yield and quality.The bud stage is one of the most vulnerable stages in the plant life cycle,and improving salt and alkali tolerance during the bud stage is particularly important to ensure the healthy growth of crops throughout the entire growth cycle.This study focuses on exploring the alleviation mechanism of melatonin on sprouting red beans under salt alkali stress,using the red bean variety Longxiaodou 5 as the experimental material.The experiment was conducted with distilled water as the control(CK)and three treatment groups,namely SA(mixed saline alkali solution),MT(melatonin solution),and MT+SA(melatonin solution with a concentration of 100 μmol/L added to SA).Compared with SA,MT+SA treatment can significantly promote the growth of red bean sprouts.The antioxidant enzyme activity,soluble protein,and soluble sugar content of red beans were significantly enhanced.The content of ROS,MDA,and H2O2 of red beans were significantly reduced.In terms of transcriptome analysis,a comparison between SA and CK,and a comparison between MT+SA and SA,resulted in 9 111 and 4 662 differentially expressed genes(DEGs),respectively.There were 4 624 upregulated genes and 4 487 downregulated genes in the SA vs CK treatment group;there were 1 800 upregulated genes and 2 862 downregulated genes in the MT+SA and SA treatment groups.GO enrichment analysis of differentially expressed genes revealed that they were mainly enriched in metabolic processes,biological regulation,transcriptional regulation activity,transport activity,catalytic activity,and other aspects;the main pathways identified through KEGG enrichment analysis were map00940,map00941,and map00943.The differential genes identified in these three pathways were validated by qRT-PCR,and the results showed that the expression trend of qRT-PCR was consistent with that of RNA-seq,thus confirming the reliability of RNA-seq data.In summary,based on phenotype,physiology,transcriptome data,and qRT-PCR data,this study found that melatonin has a significant effect on the resistance of sprouting red beans to salt alkali stress.
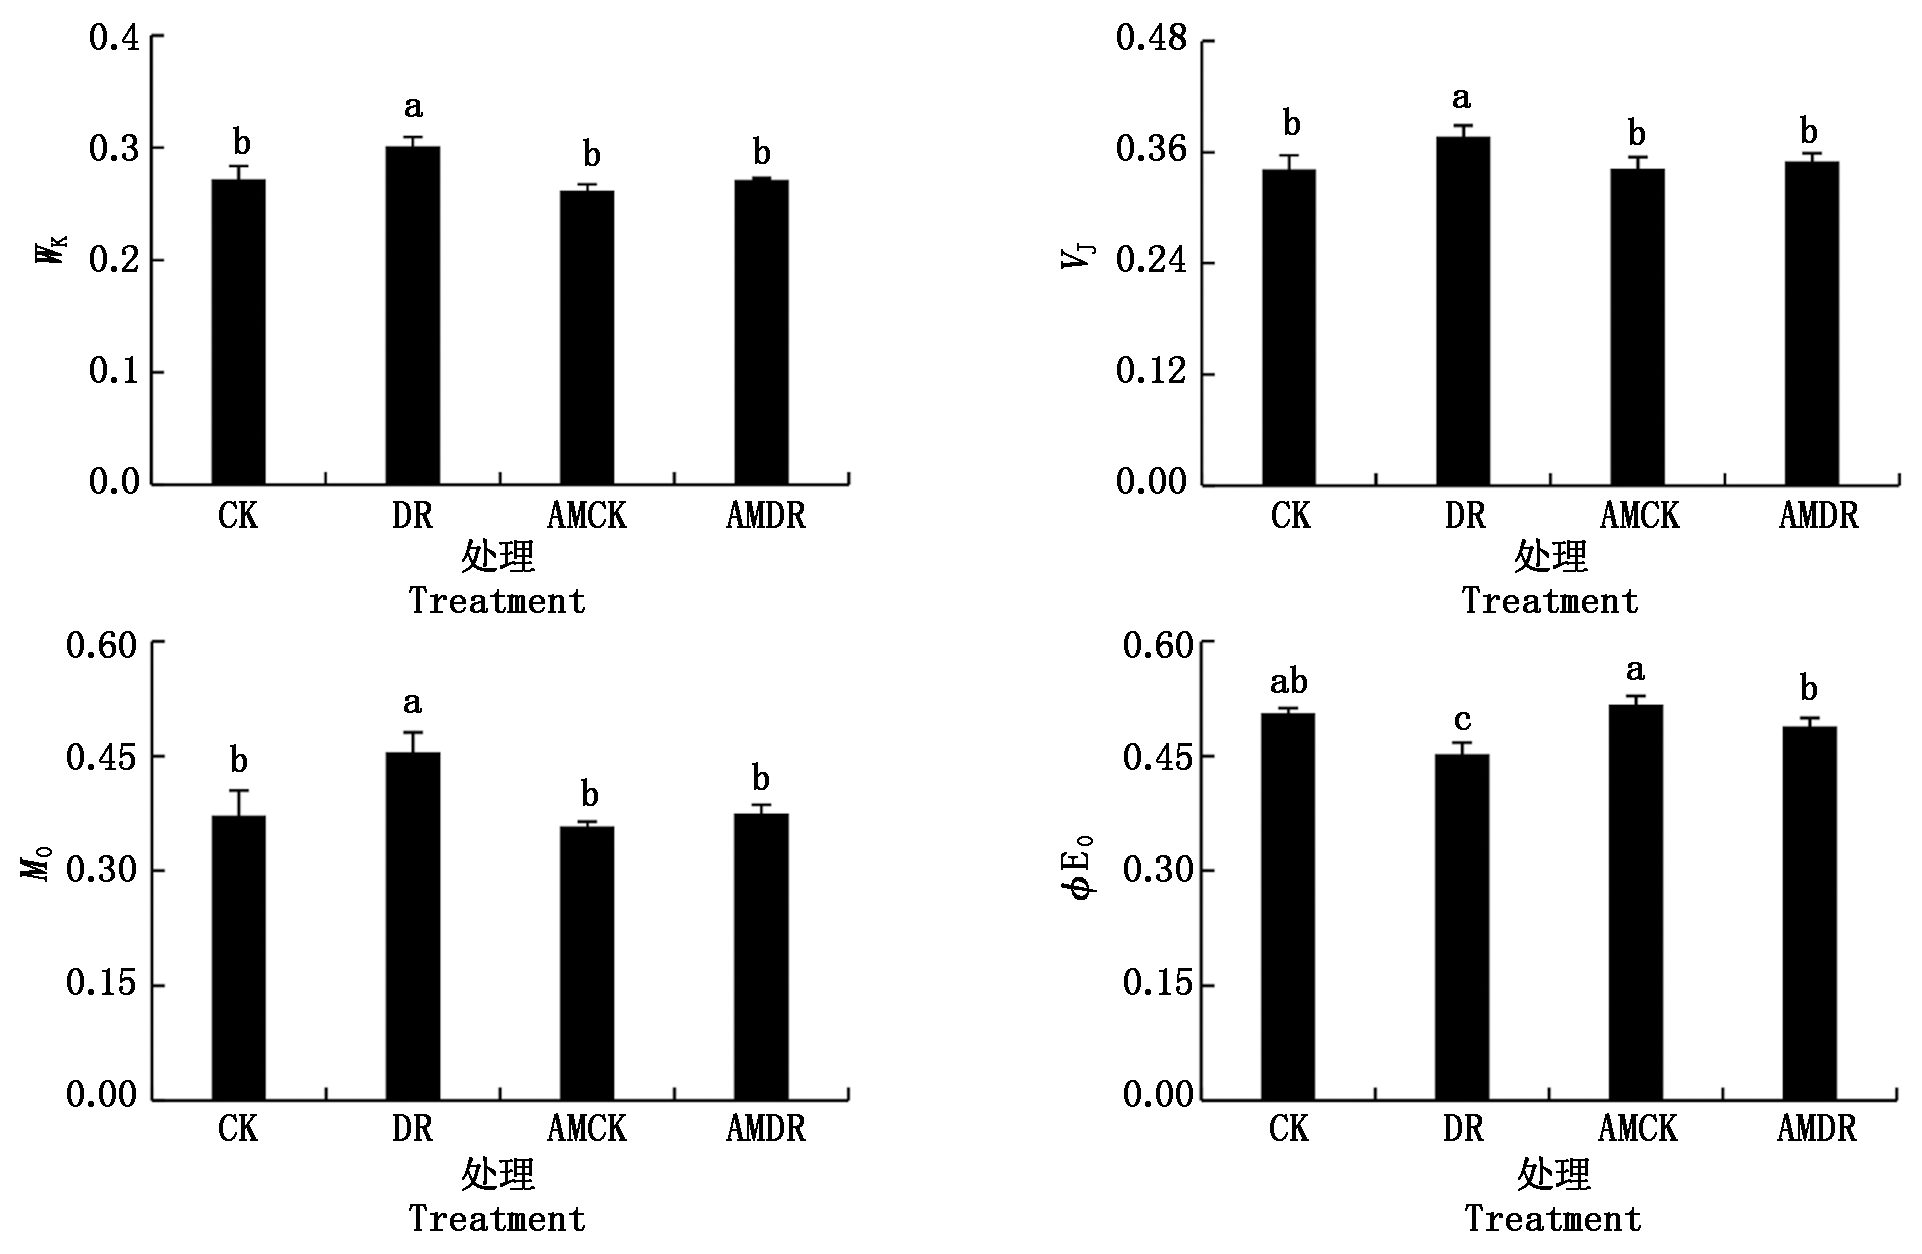
This study investigated the effects of arbuscular mycorrhizal fungi(AMF)inoculation on the physiological characteristics and the accumulation of active components in the medicinal tree peony Paeonia ostii Fengdan under drought stress,with the aim of providing a theoretical basis for utilizing AMF to enhance the drought resistance and medicinal quality of this species.Using P.ostii seedlings as experimental material,four treatments were established:well-watered control(CK),drought stress(DR),AMF inoculation under well-watered conditions(AMCK),and AMF inoculation under drought stress(AMDR).Indicators related to plant redox homeostasis(including superoxide dismutase(SOD)activity,proline and soluble sugar content),photosynthetic characteristics(including photosynthetic pigment content,gas exchange parameters,and chlorophyll fluorescence parameters),and the content of active components(such as total flavonoids,paeonol,and paeoniflorin)in the root bark were measured.The results showed that drought stress induced significant accumulation of malondialdehyde(MDA)and reactive oxygen species(ROS),with increases of 33.53% and 50.04%, respectively,while photosynthetic performance was markedly inhibited.This was specifically evidenced by significant decreases in net photosynthetic rate(Pn),stomatal conductance(Gs),transpiration rate(Tr),maximum photochemical efficiency(Fv/Fm),and photosynthetic performance index(PIabs).AMF inoculation effectively alleviated these adverse effects.It was found that inoculation not only significantly increased SOD activity by 6.36% and elevated the contents of osmoregulatory substances (soluble proteins, soluble sugars, and proline) by 23.72%, 53.31%, and 14.70%, respectively,but also reduced ROS levels,thereby enhancing cell membrane stability.A significant recovery was also observed in photosynthetic pigment content and photosynthetic and chlorophyll fluorescence parameters(Pn,Gs,Tr,Fv/Fm,and PIabs).Furthermore,under drought stress conditions,AMF inoculation further promoted the accumulation of secondary metabolites in the root bark,including total flavonoids,paeonol,and paeoniflorin,with increases of 18.67%, 11.39%, and 5.17%, respectively, compared with the DR treatment.In conclusion,it is demonstrated that AMF can effectively mitigate the physiological damage caused by drought stress in P.ostii by enhancing its antioxidant capacity,promoting the accumulation of osmoregulatory substances,and improving photosynthetic performance.Simultaneously,AMF boosts the content of secondary metabolites in the root bark.
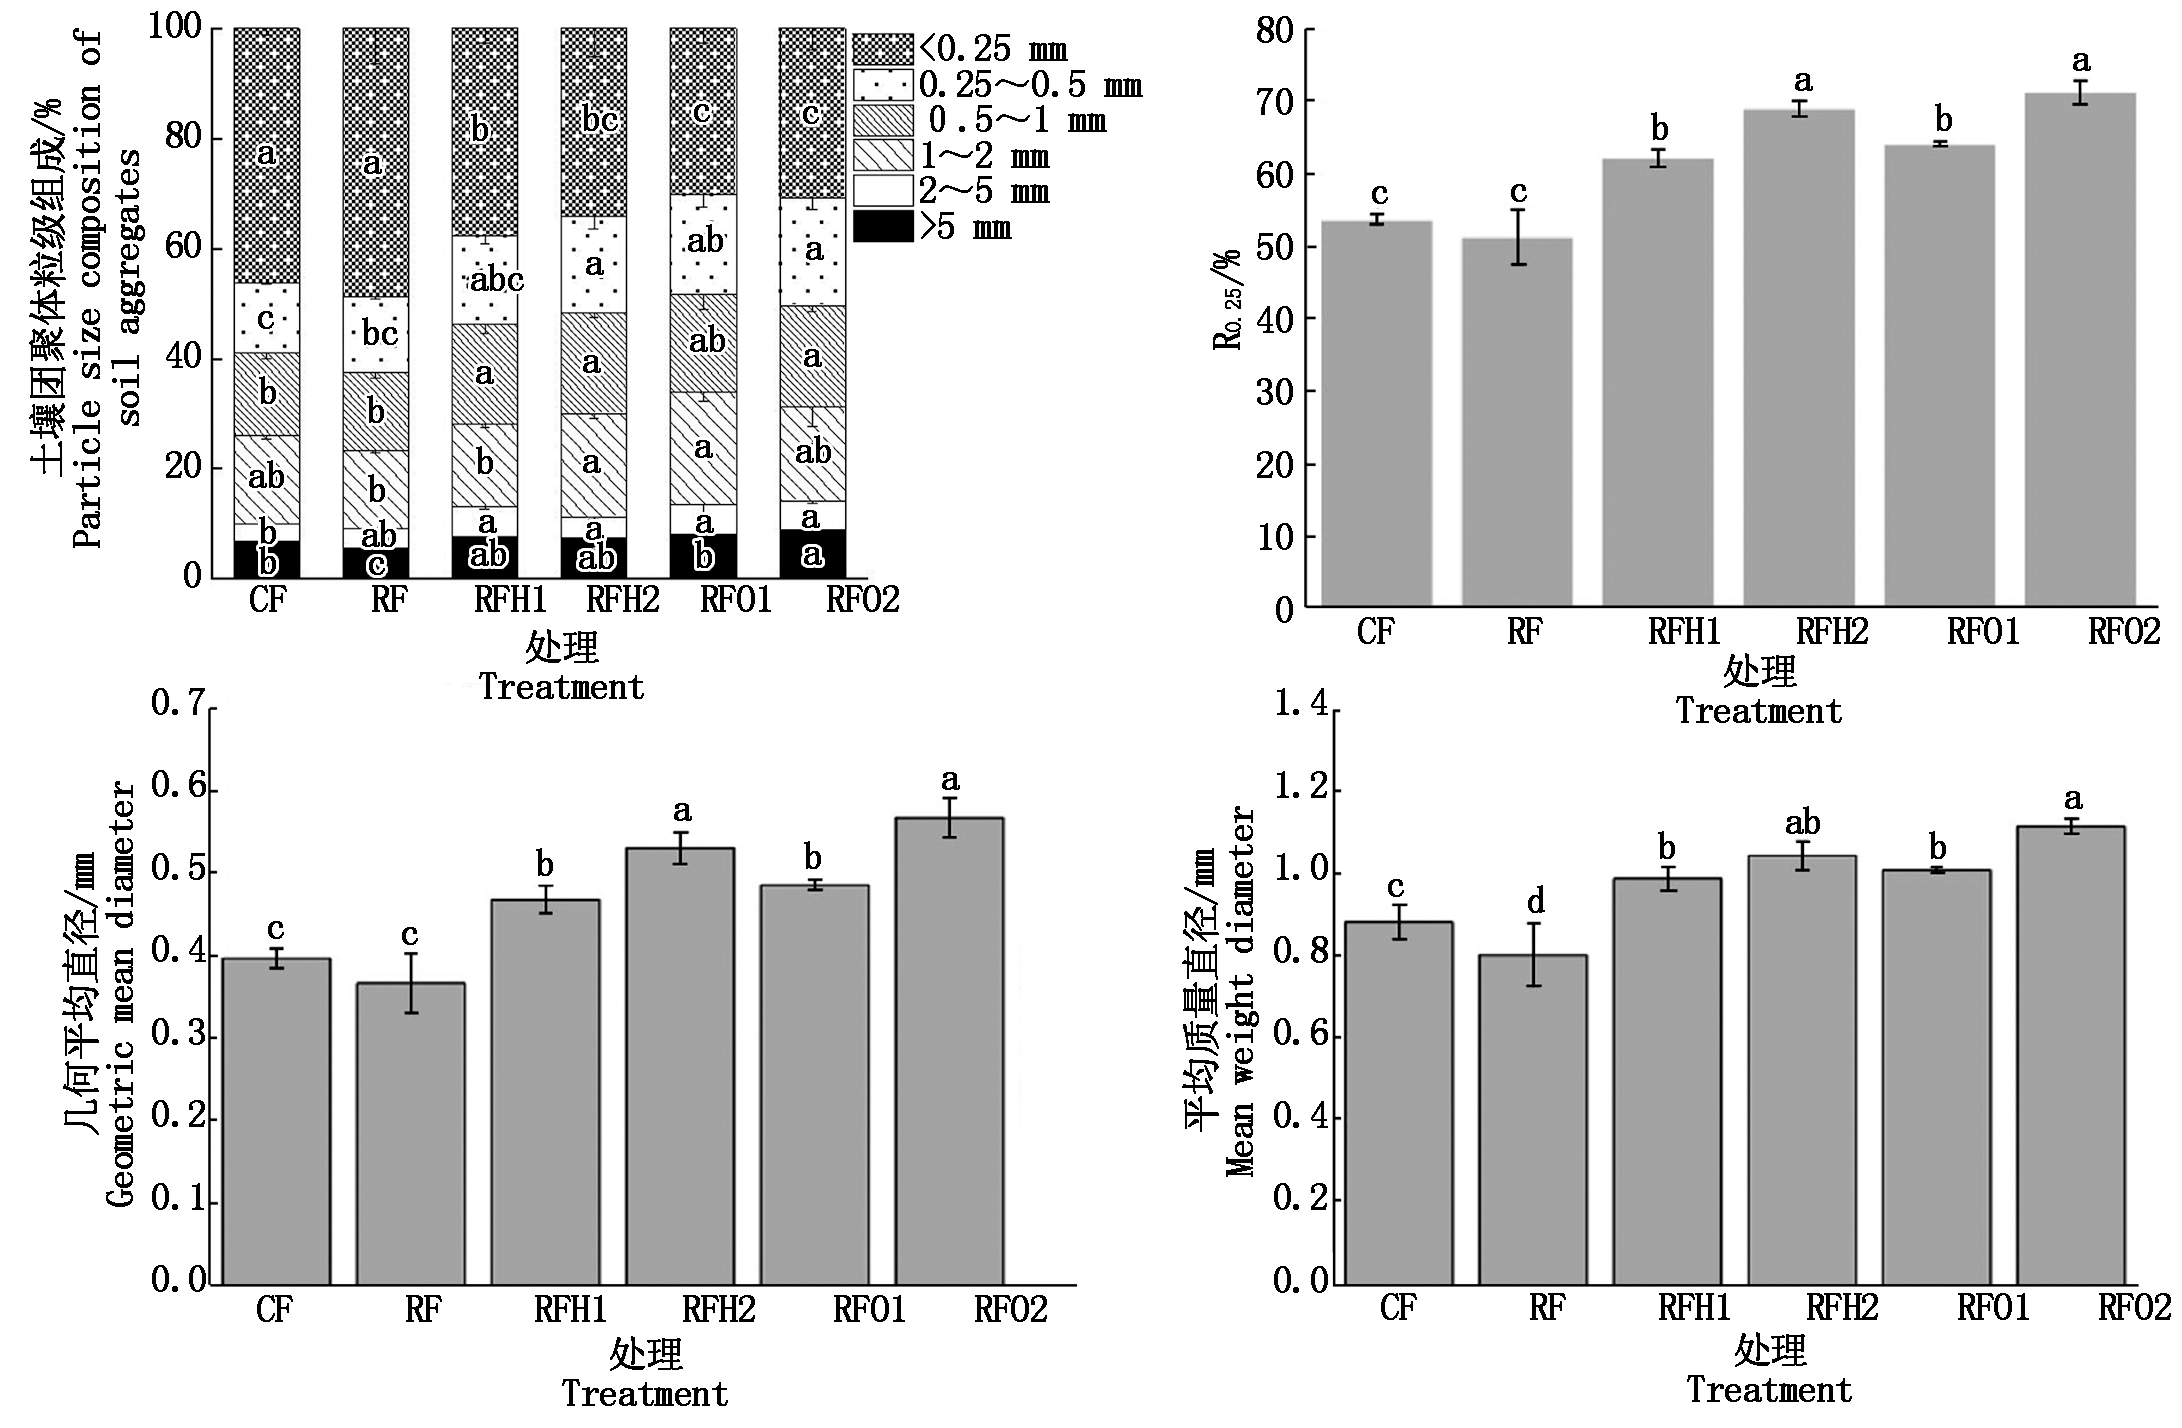
To elucidate the intrinsic mechanisms underlying the amelioration of degraded black soils by organic fertilizers,and thereby provide a scientific basis for optimizing chemical fertilizer reduction strategies while safeguarding long-term soil productivity in the target region,a maize field trial was conducted in 2024.Six fertilization treatments were established,as follows:CF(regular fertilization),RF(20% reduction of chemical fertilizer),RFH1(RF supplemented with 200 kg/ha humic acid fertilizer),RFH2(RF supplemented with 330 kg/ha humic acid fertilizer),RFO1(RF supplemented with 200 kg/ha chicken manure organic fertilizer)and RFO2(RF supplemented with 330 kg/ha chicken manure organic fertilizer).Compared to CF and RF,the RFH2,RFO1,and RFO2 treatments significantly reduced soil bulk density by 12.4%—20.7%.All organic amendment treatments improved soil structure stability,and the fraction of water-stable aggregates(>0.25 mm)increased by 15.9%—39.1%,while soil pH decreased by 0.8%—1.2%.RFO1 and RFO2 elevated soil alkaline hydrolyzable nitrogen content by 14.6%—28.5%.Additionally,every organic treatment increased soil available phosphorus and soil organic matter contents by 8.7%—36.8% and 11.0%—48.9%,respectively,with RFO2 showing the most pronounced improvements.At the same time,RFH2,RFO1,and RFO2 significantly enhanced the activities of key soil enzymes(urease,amylase,catalase,alkaline phosphatase,and sucrase)by 18.1%—72.4%.In terms of grain yield,RFH2,RFO1,and RFO2 outperformed CF by 0.5%—13.9%,and all organic amended treatments increased by 16.7%—36.6% compared with RF,with RFO2 achieving the highest yield.Organic amended treatments also improved partial productivity and agronomic efficiency of fertilizers by 11.9%—31.1%,while RFO2 exhibited 25.8%—36.6% higher nitrogen and phosphorus partial productivity compared to CF and RF.Overall,chemical fertilizer reduction combined with organic fertilizer effectively improved soil physicochemical properties,enhanced soil enzyme activities,and increased nutrient-use efficiency.This practice not only maintained maize yield but also contributed to the restoration of degraded black soils.Thus,20% chemical fertilizer reduction combined with 330 kg/ha chicken manure organic fertilizer(RFO2)was identified as the optimal fertilization strategy for degraded black soils in this experiment.
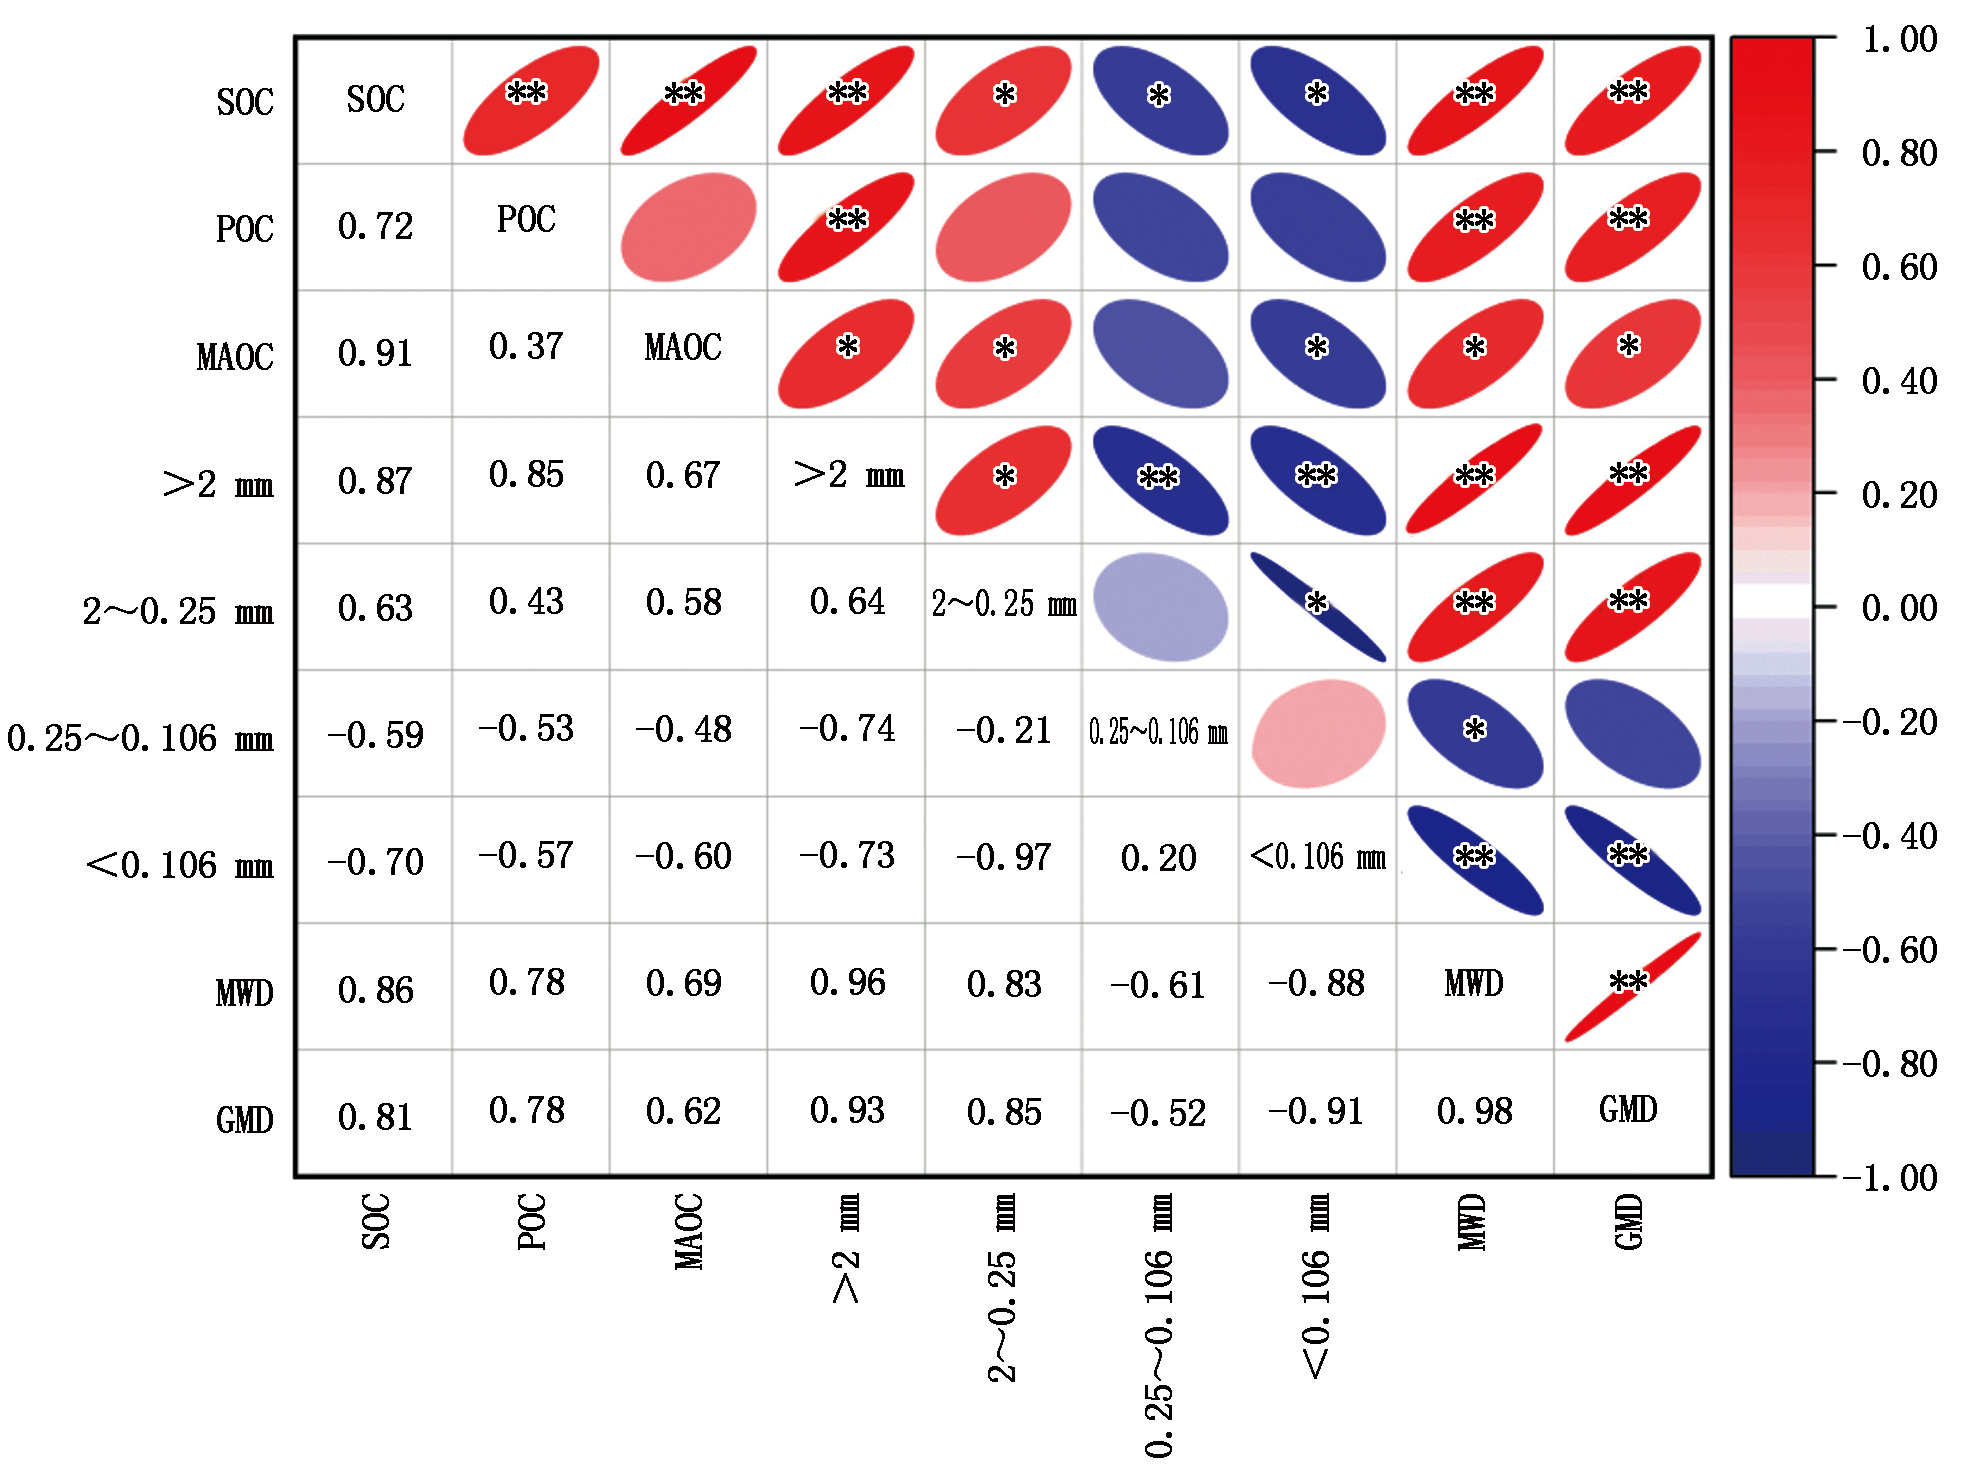
In order to explore the effects of mulching measures(such as straw returning and plastic film mulching)on soil aggregates and organic carbon in semi-arid rain-fed agricultural areas.The potato-wheat rotation system in the dryland of Longzhong was used as the research object.The effects of straw stubble mulching(SR),straw strip mulching(SM),plastic film mulching(PM)and open field(CK)on soil organic carbon components(SOC,POC,MAOC),aggregate stability and aggregate carbon contribution rate were systematically analyzed.By evaluating the soil improvement effect of different mulching measures,integrating the intrinsic indicators such as soil organic carbon component dynamics,aggregate stability and crop productivity and water use efficiency data reported in the literature,the optimal mulching mode that can synergistically improve organic carbon and aggregate stability was screened.The results showed that the organic carbon content and aggregate stability of potato and wheat were improved by three mulching measures in potato-wheat rotation,and the effect of SR treatment was the best.Compared with CK treatment,the SOC content of SR and SM treatment increased by 15.38% and 7.78%,and the SOC content of PM treatment did not change significantly.SR and SM treatment increased the POC content in 0—20 cm soil layer by 18.54%,16.62%,and PM decreased the POC content in 0—20 cm soil layer by 3.06%.The overall trend of MAOC increased in each treatment was SR>SM>PM>CK.Compared with CK treatment,SR and SM treatments increased the proportion of MWD,GMD,>2 mm aggregates and SOC content,while PM significantly reduced MWD and GMD.SR increased the SOC enrichment coefficient of>2 mm aggregates by 17.26% in 0—20 cm soil layer,and SR and SM treatments significantly increased the SOC contribution rate of>2 mm aggregates.In summary,under the conditions of this experiment,all three mulching measures can increase soil organic carbon content,and straw stubble mulching returning to the field has the best effect in improving soil organic carbon content and aggregate stability,effectively improving soil fertility.
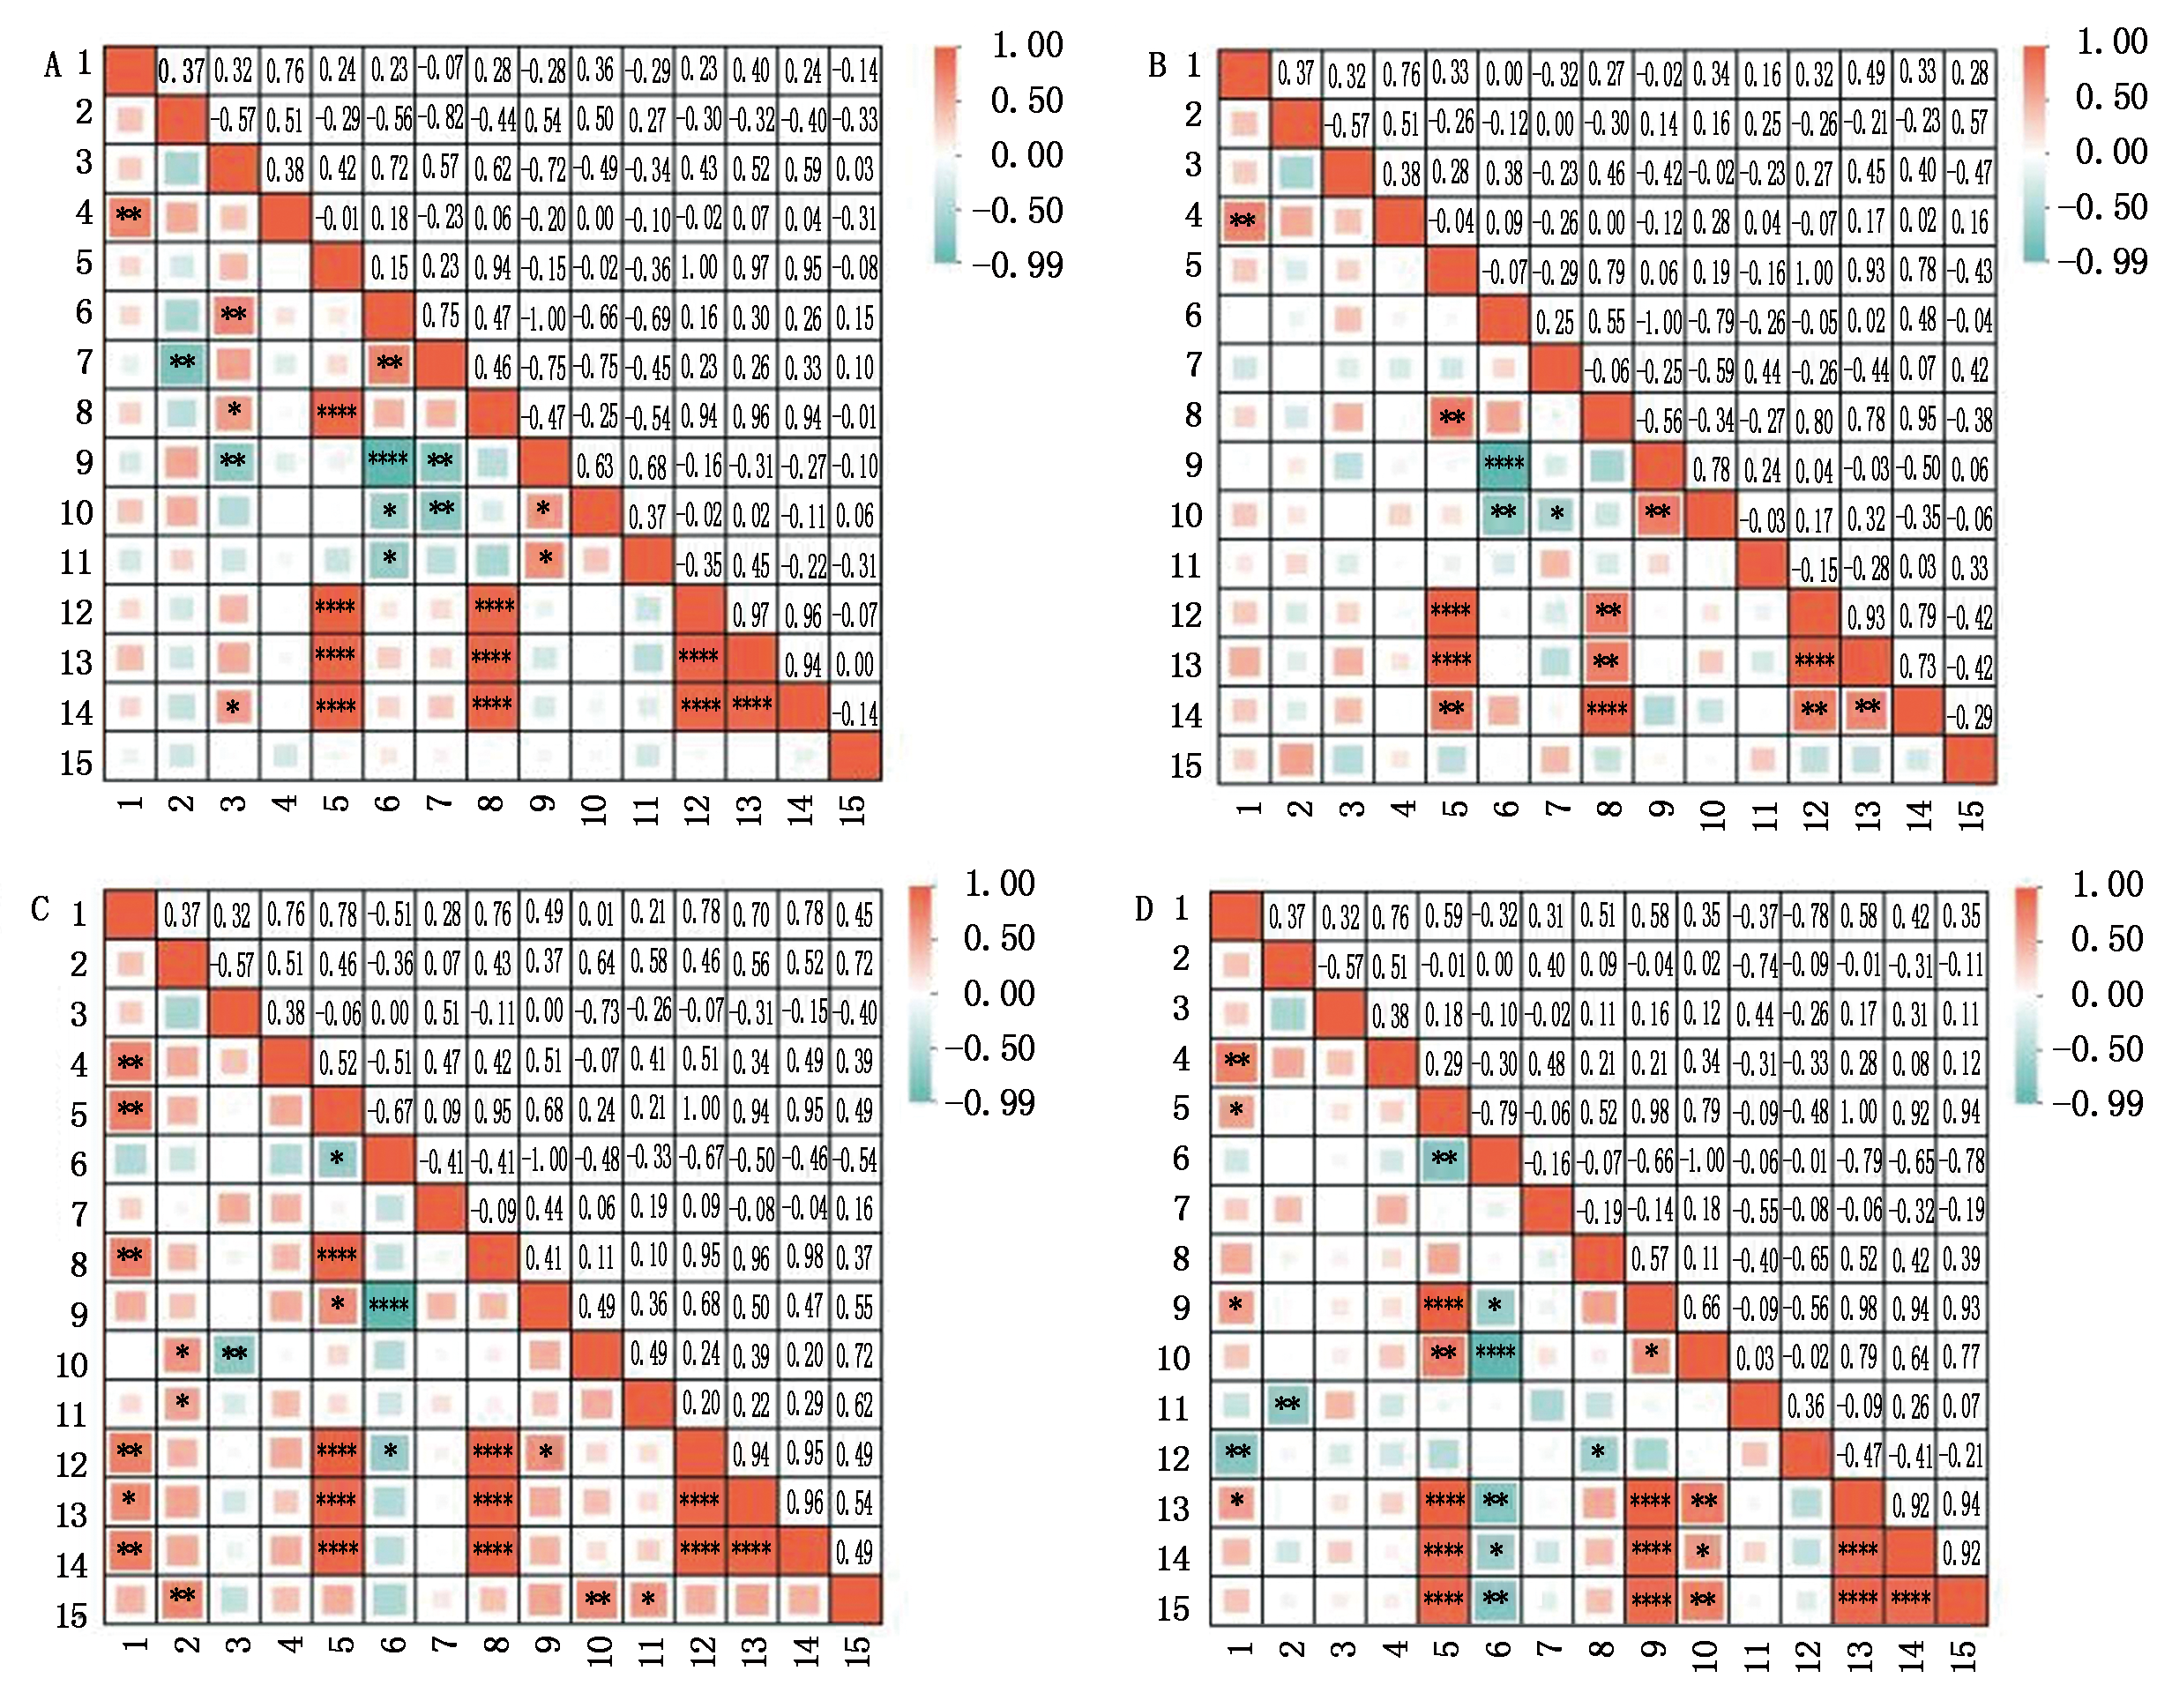
To clarify the effects of different nitrogen forms combined with magnesium fertilization on potato in the northern foothills of Yinshan Mountains,a field experiment was conducted in Hohhot,Inner Mongolia.The experiment included two nitrogen forms(nitrate nitrogen and ammonium nitrogen)and two magnesium application rates(0,45 kg/ha),resulting in four treatments.The impacts of these treatments on potato yield and its components,dry matter,nitrogen,magnesium,and potassium contents,their accumulation,and nitrogen use efficiency throughout the growth period were analyzed.The results showed that,compared with the application of nitrate nitrogen alone,the combined application of nitrate nitrogen and magnesium significantly increased potato leaf dry matter(21.24%—75.60%)and tuber dry matter(22.04%—62.36%)from tuber initiation stage to harvest stage,as well as whole-plant dry matter(32.54%—70.28%)from seedling stage to harvest stage.It also enhanced nitrogen accumulation(44.01%—69.84%)in tuber initiation stage,starch accumulation stage and harvest stage,whole-plant magnesium accumulation(37.66%—84.21%)during the entire growth period,whole-plant potassium accumulation (24.30%—54.95%) from tuber initiation stage to harvest stage, nitrogen uptake efficiency (30.77%—66.67%) during the entire growth period,and partial factor productivity of nitrogen fertilizer at harvest stage(35.13%).Correlation analysis indicated that potato dry matter,whole-plant nitrogen,magnesium,and potassium accumulation were significantly positively correlated with nitrogen uptake efficiency during the entire growth period(r was all greater than 0.78).At harvest stage,whole-plant potassium content showed a highly significant negative correlation with yield(r was -0.78).Principal component analysis further demonstrated that the total variation among treatments during the entire growth period was highly dispersed,indicating significant differences between nitrogen form combined with magnesium fertilization treatments and those without magnesium application.These findings provide a new strategy for improving nitrogen use efficiency and optimizing nitrogen-magnesium management in potato production in the northern foothills of Yinshan Mountains.
Soil salinization is an important constraint on the improvement of farmland productivity.In view of the poor quality and low productivity of coastal salinized soil,a pot experiment was set up to explore the effects of different dosages and ratios of organic soil conditioners on the growth and soil properties of soybean in coastal salinized farmland.The results showed that the comprehensive analysis of the physiological indicators of salt resistance,such as SOD,CAT,and MDA,showed that T3(lactose peptide 225 kg/ha,lactose peptide:inositol=10:1)and T5(lactose peptide 225 kg/ha,lactose peptide:inositol=20:1)had good improvement effects.Comparing the growth trend in the early and medium stages,T5 treatment showed good growth status,followed by T2(lactose peptide dosage 225 kg/ha,lactose peptide:inositol=5:1).In terms of dry matter accumulation and nutrient uptake,T16(lactose peptide dosage 900 kg/ha,lactose peptide:inositol=15:1),T17(lactose peptide dosage 900 kg/ha,lactose peptide:inositol=20:1)and T2 were better than those of the control treatment.The partial productivity and agronomic efficiency were the highest in T2,which were 1.11,0.87 g/kg,and the compound ratio was 5:1—15:1,which had a better effect on improving soil water-soluble ions.Taken together,the T2 and T3 treatments with a lactose peptide dosage of 225 kg/ha and a lactose peptide and inositol compound ratio of 5:1~10:1 were the best in terms of saving cost and maintaining the improvement effect.
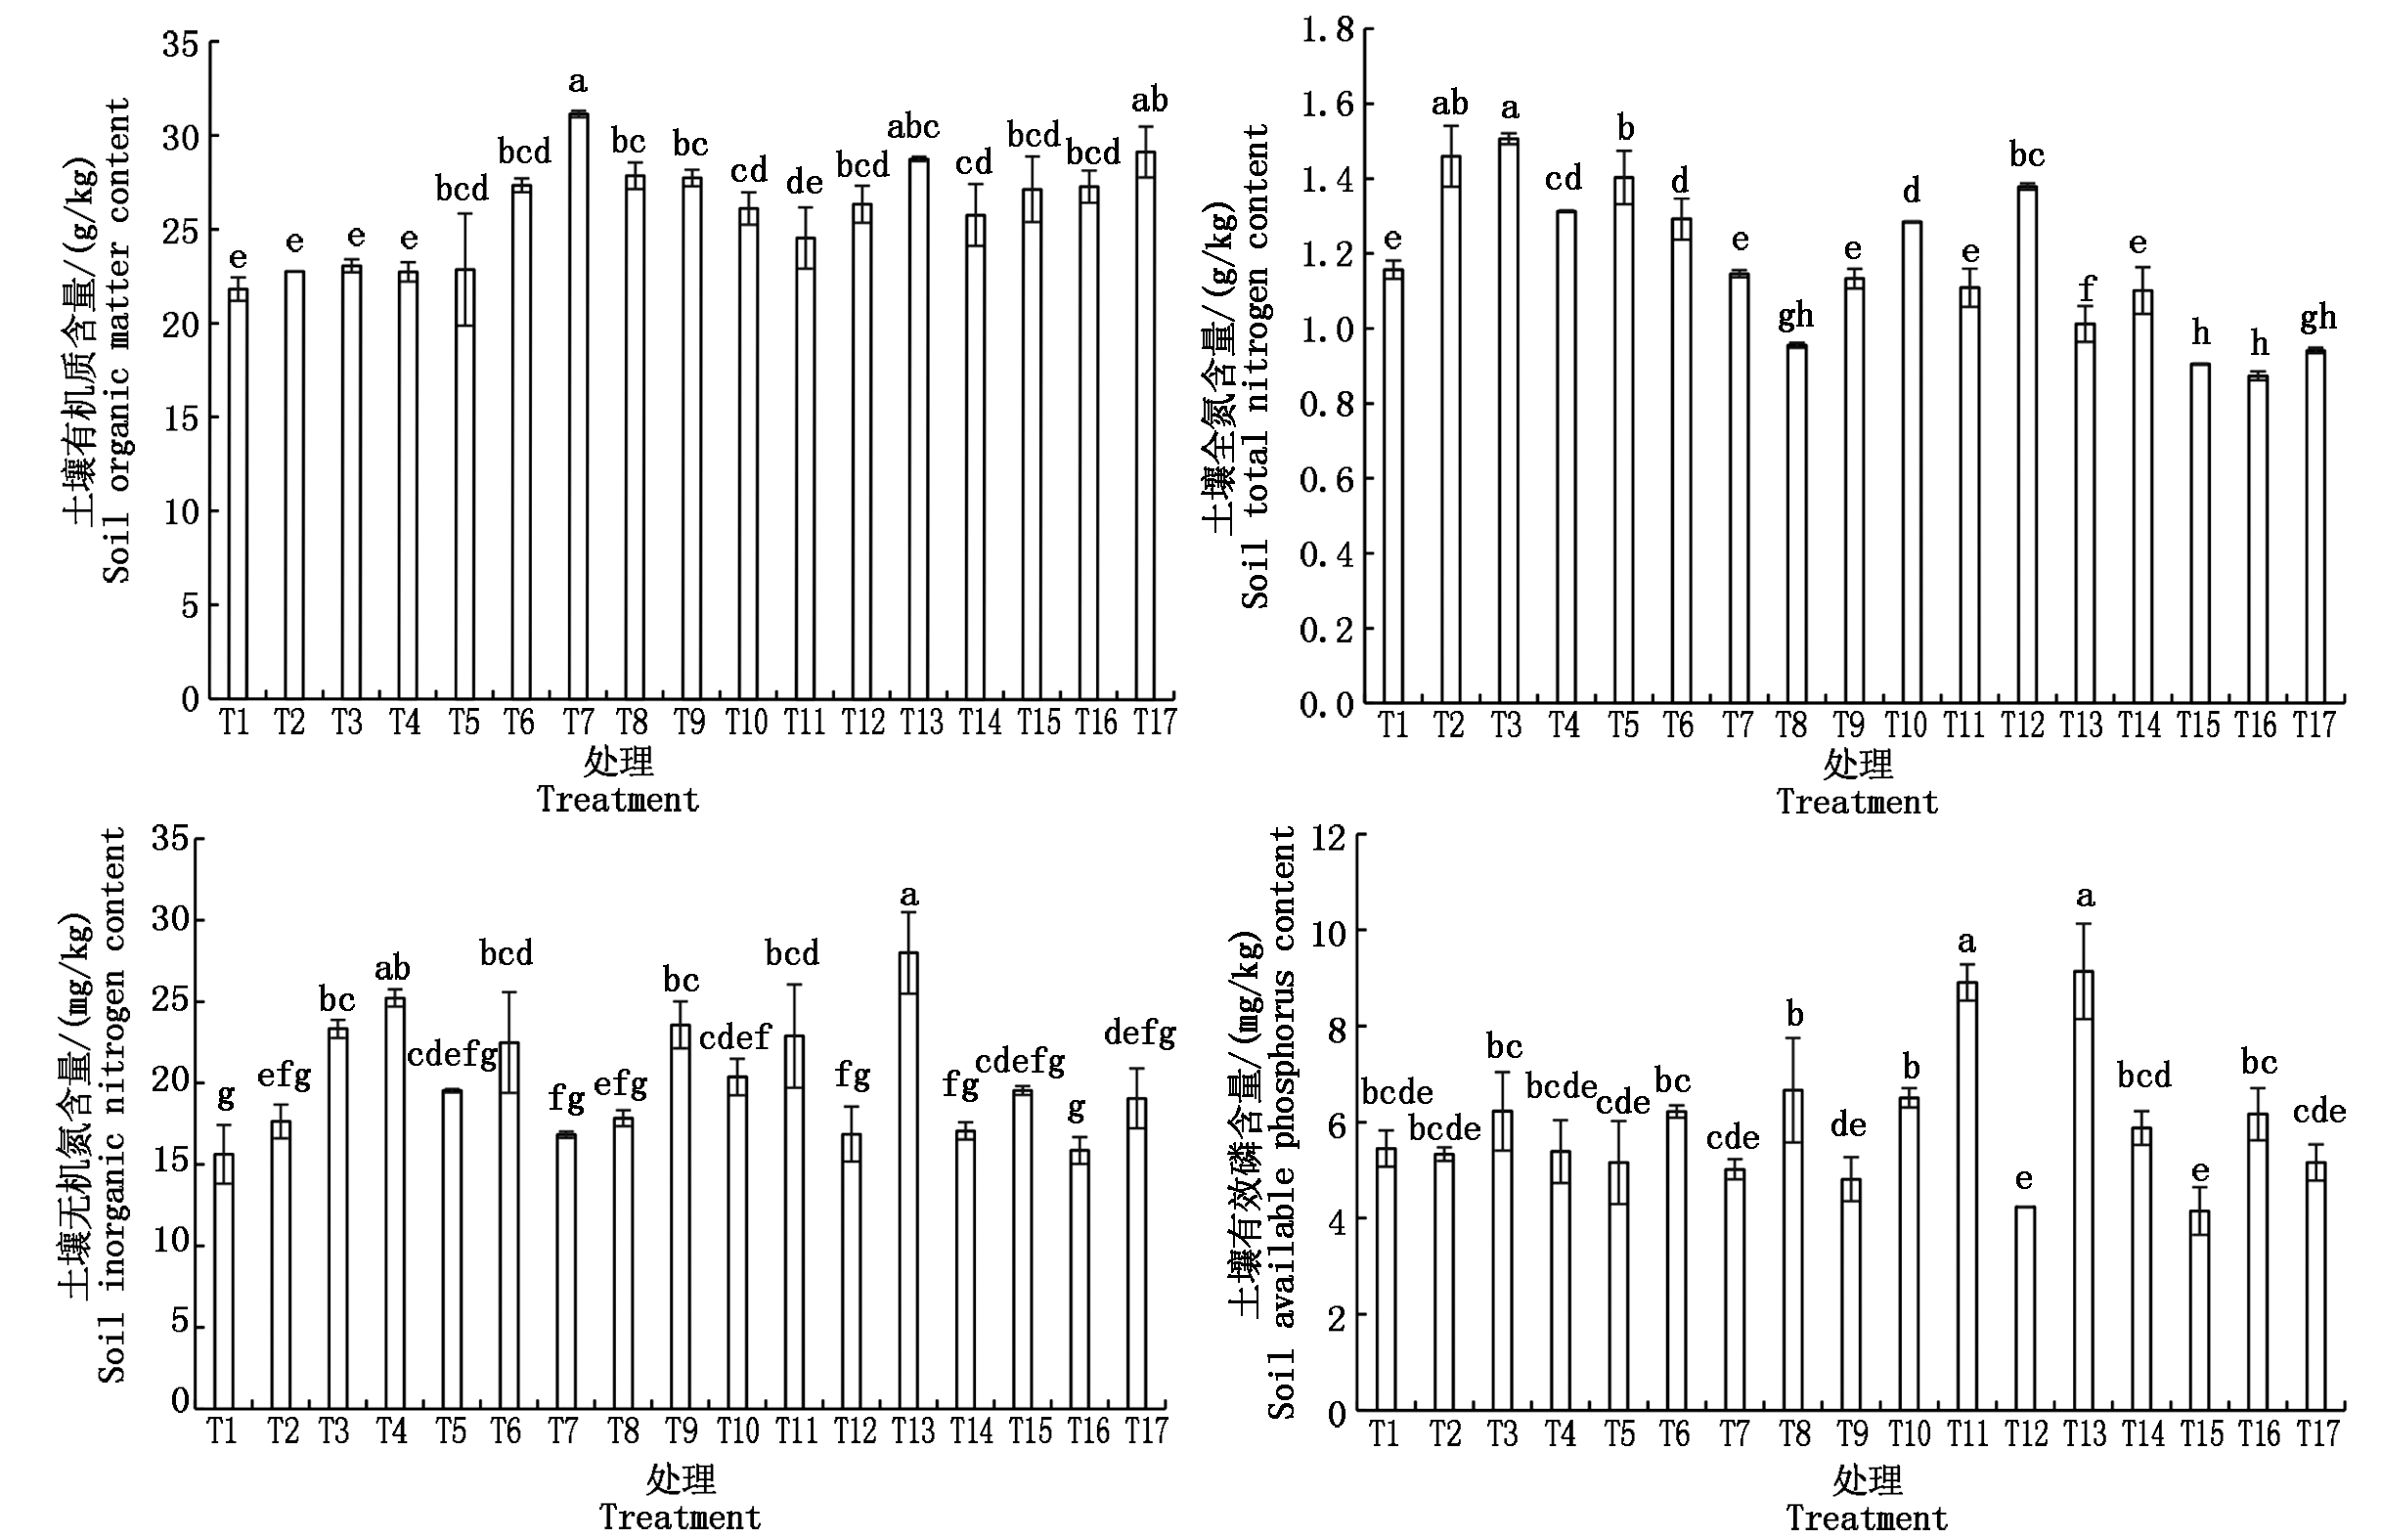
To investigate the effects of partially substituting chemical fertilizer with biochar-based organic fertilizer derived from cattle manure on N2O emissions in summer maize fields and the underlying regulatory mechanisms,a field plot experiment was conducted from 2023 to 2024 at the Dishang Experimental Station of the Institute of Cereal and Oil Crops,Hebei Academy of Agriculture and Forestry Sciences.Four fertilization treatments were set up:CF(conventional nitrogen-phosphorus-potassium fertilizer),and COF1,COF2,and COF3,which replaced 15%,30%,and 45% of the chemical nitrogen in CF with an equivalent amount of nitrogen from biochar-based organic fertilizer derived from cattle manure,respectively.The study analyzed the effects of different fertilization regimes on N2O emissions,plant nitrogen uptake,soil physicochemical properties,abundance of nitrogen-cycle-related functional genes,and grain yield.The results showed that:(1)Compared with CF,the cumulative N2O emissions under COF1,COF2,and COF3 treatments were significantly reduced by 39.4%,42.5%,and 43.8%,respectively.While reducing emissions,COF1 and COF2 increased plant nitrogen uptake by 45.9% and 24.5%,and yield by 10.6% and 11.7%,respectively.(2)COF1 and COF2 significantly promoted the formation of water-stable aggregates(WSA)(increased by 12.7% and 16.5% compared with CF,respectively)and reduced the abundance of key denitrification functional genes(norB,norC,narH),while enhancing the expression of genes related to dissimilatory nitrate reduction to ammonium(DNRA)(napA,napB),thereby promoting the accumulation of soil nitrate and ammonium nitrogen.(3)COF1 and COF2 increased root biomass and the activities of nitrate reductase and glutamine synthetase in roots and leaves,synergistically improving plant nitrogen uptake and utilization.Correlation analysis further revealed that soil WSA formation was significantly negatively correlated with the abundance of key denitrification genes(norB/C,narH),while positively correlated with plant nitrogen uptake,root biomass,and nitrogen metabolism enzyme activities.In conclusion,replacing 15%—30% of chemical fertilizer with biochar-based organic fertilizer derived from cattle manure can directly inhibit the denitrification process and reduce N2O emissions by promoting the formation of water-stable soil aggregates.Simultaneously,it indirectly reduces N2O emissions by enhancing the DNRA pathway and plant nitrogen uptake capacity,which competes with denitrification for nitrate substrates.
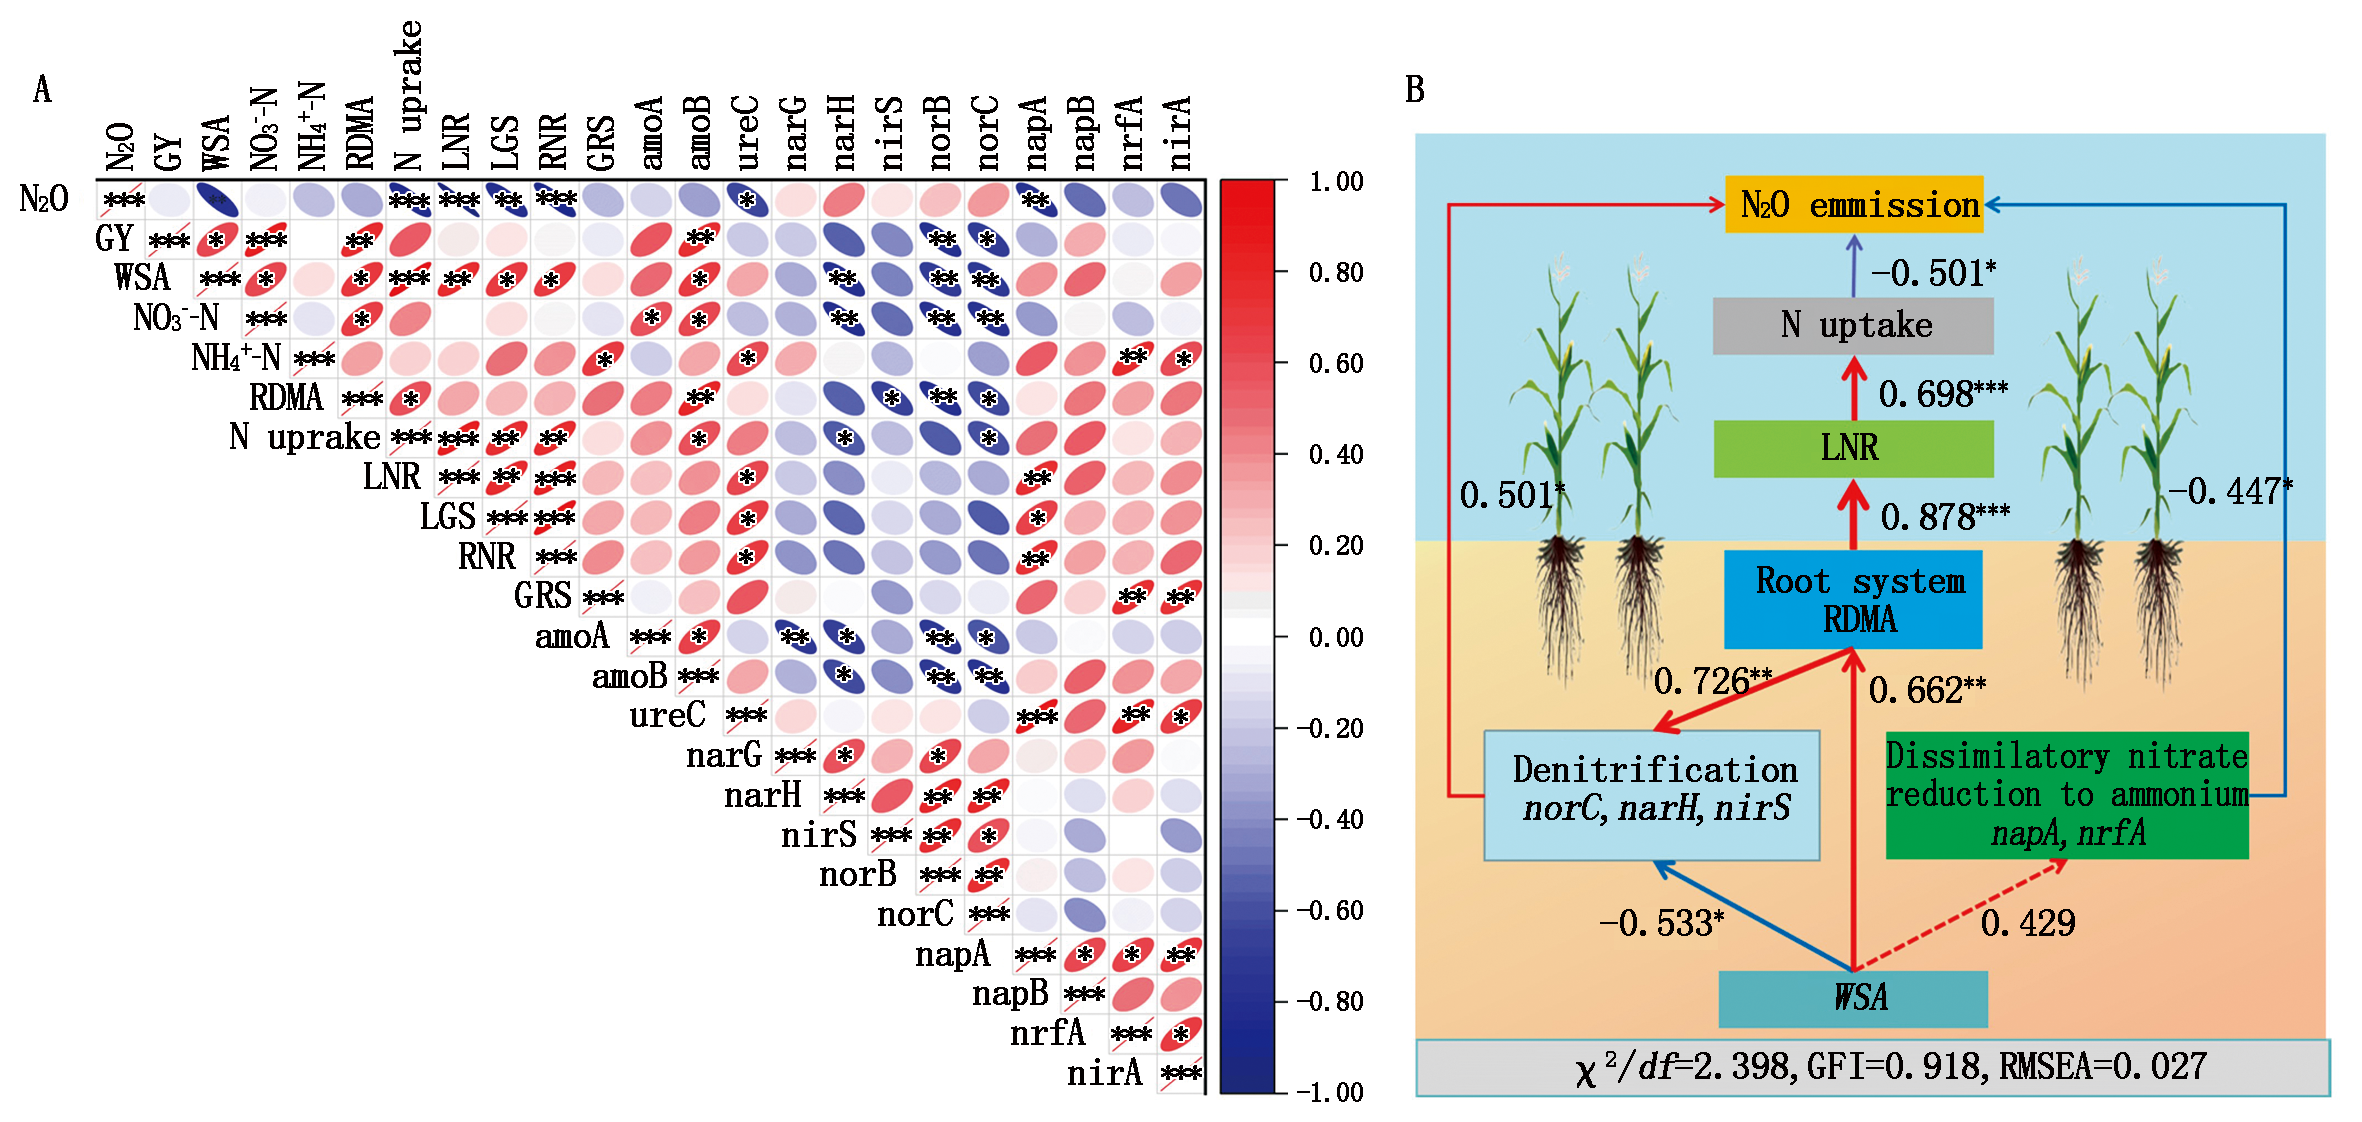
To elucidate the regulatory effects of water-fertilizer coupling on fruit quality and taste substance accumulation in flavor tomatoes,greenhouse experiments were conducted using Yutianyou 6(premium flavor type)and 2442(common type)as test materials.Six water-fertilizer coupling treatments were established(W1F1:70%—75% ETc+180-90-150 kg/ha;W1F2:70%—75% ETc+240-120-200 kg/ha;W1F3:70%—75% ETc+300-150-250 kg/ha;W2F1:80%—85% ETc+180-90-150 kg/ha;W2F2:80%—85% ETc+240-120-200 kg/ha;W2F3:80%—85% ETc+300-150-250 kg/ha).The effects on yield,nutritional quality,sugar-acid components,and dynamic accumulation of soluble solids in greenhouse tomatoes were investigated.The results demonstrated that: both irrigation and fertilization exerted extremely significant effects on tomato yield.The W2 regime increased yields of Yutianyou 6 and 2442 by 22.3% and 19.6%,respectively,compared with W1.The highest yields were obtained under F3,reaching 33 720,33 480 kg/ha,respectively.Quality traits exhibited cultivar-specific responses.Vitamin C content was optimized in Yutianyou 6 under W1F2(0.383 mg/g)and in 2442 under W1F3(0.224 mg/g).Soluble protein accumulation was predominantly fertilizer-driven,peaking in Yutianyou 6 at W2F2(7.02 mg/g).Sugar-acid accumulation patterns varied between cultivars.Yutianyou 6 showed optimal glucose,citric acid,and malic acid accumulation under W2F2,with maximal fructose and sugar-acid ratio(8.326)achieved under W2F1.Conversely,2442 displayed weaker water responses,attaining its highest sugar-acid ratio(9.346)under W2F2.Soluble solids(Brix)displayed a rapid initial increase followed by stabilization,with W1F2 producing the optimal synergistic effect(peak 10.19% in Yutianyou 6)while conserving 12.5%—17.6% irrigation water and 20% fertilizer compared with W2F1.In conclusion,moderate water deficit combined with optimized fertilization can simultaneously enhance yield and quality.Yutianyou 6 is recommended for the high water-low fertilizer regime(W2F1),whereas 2442 is suited to the high water medium fertilizer regime(W2F2).
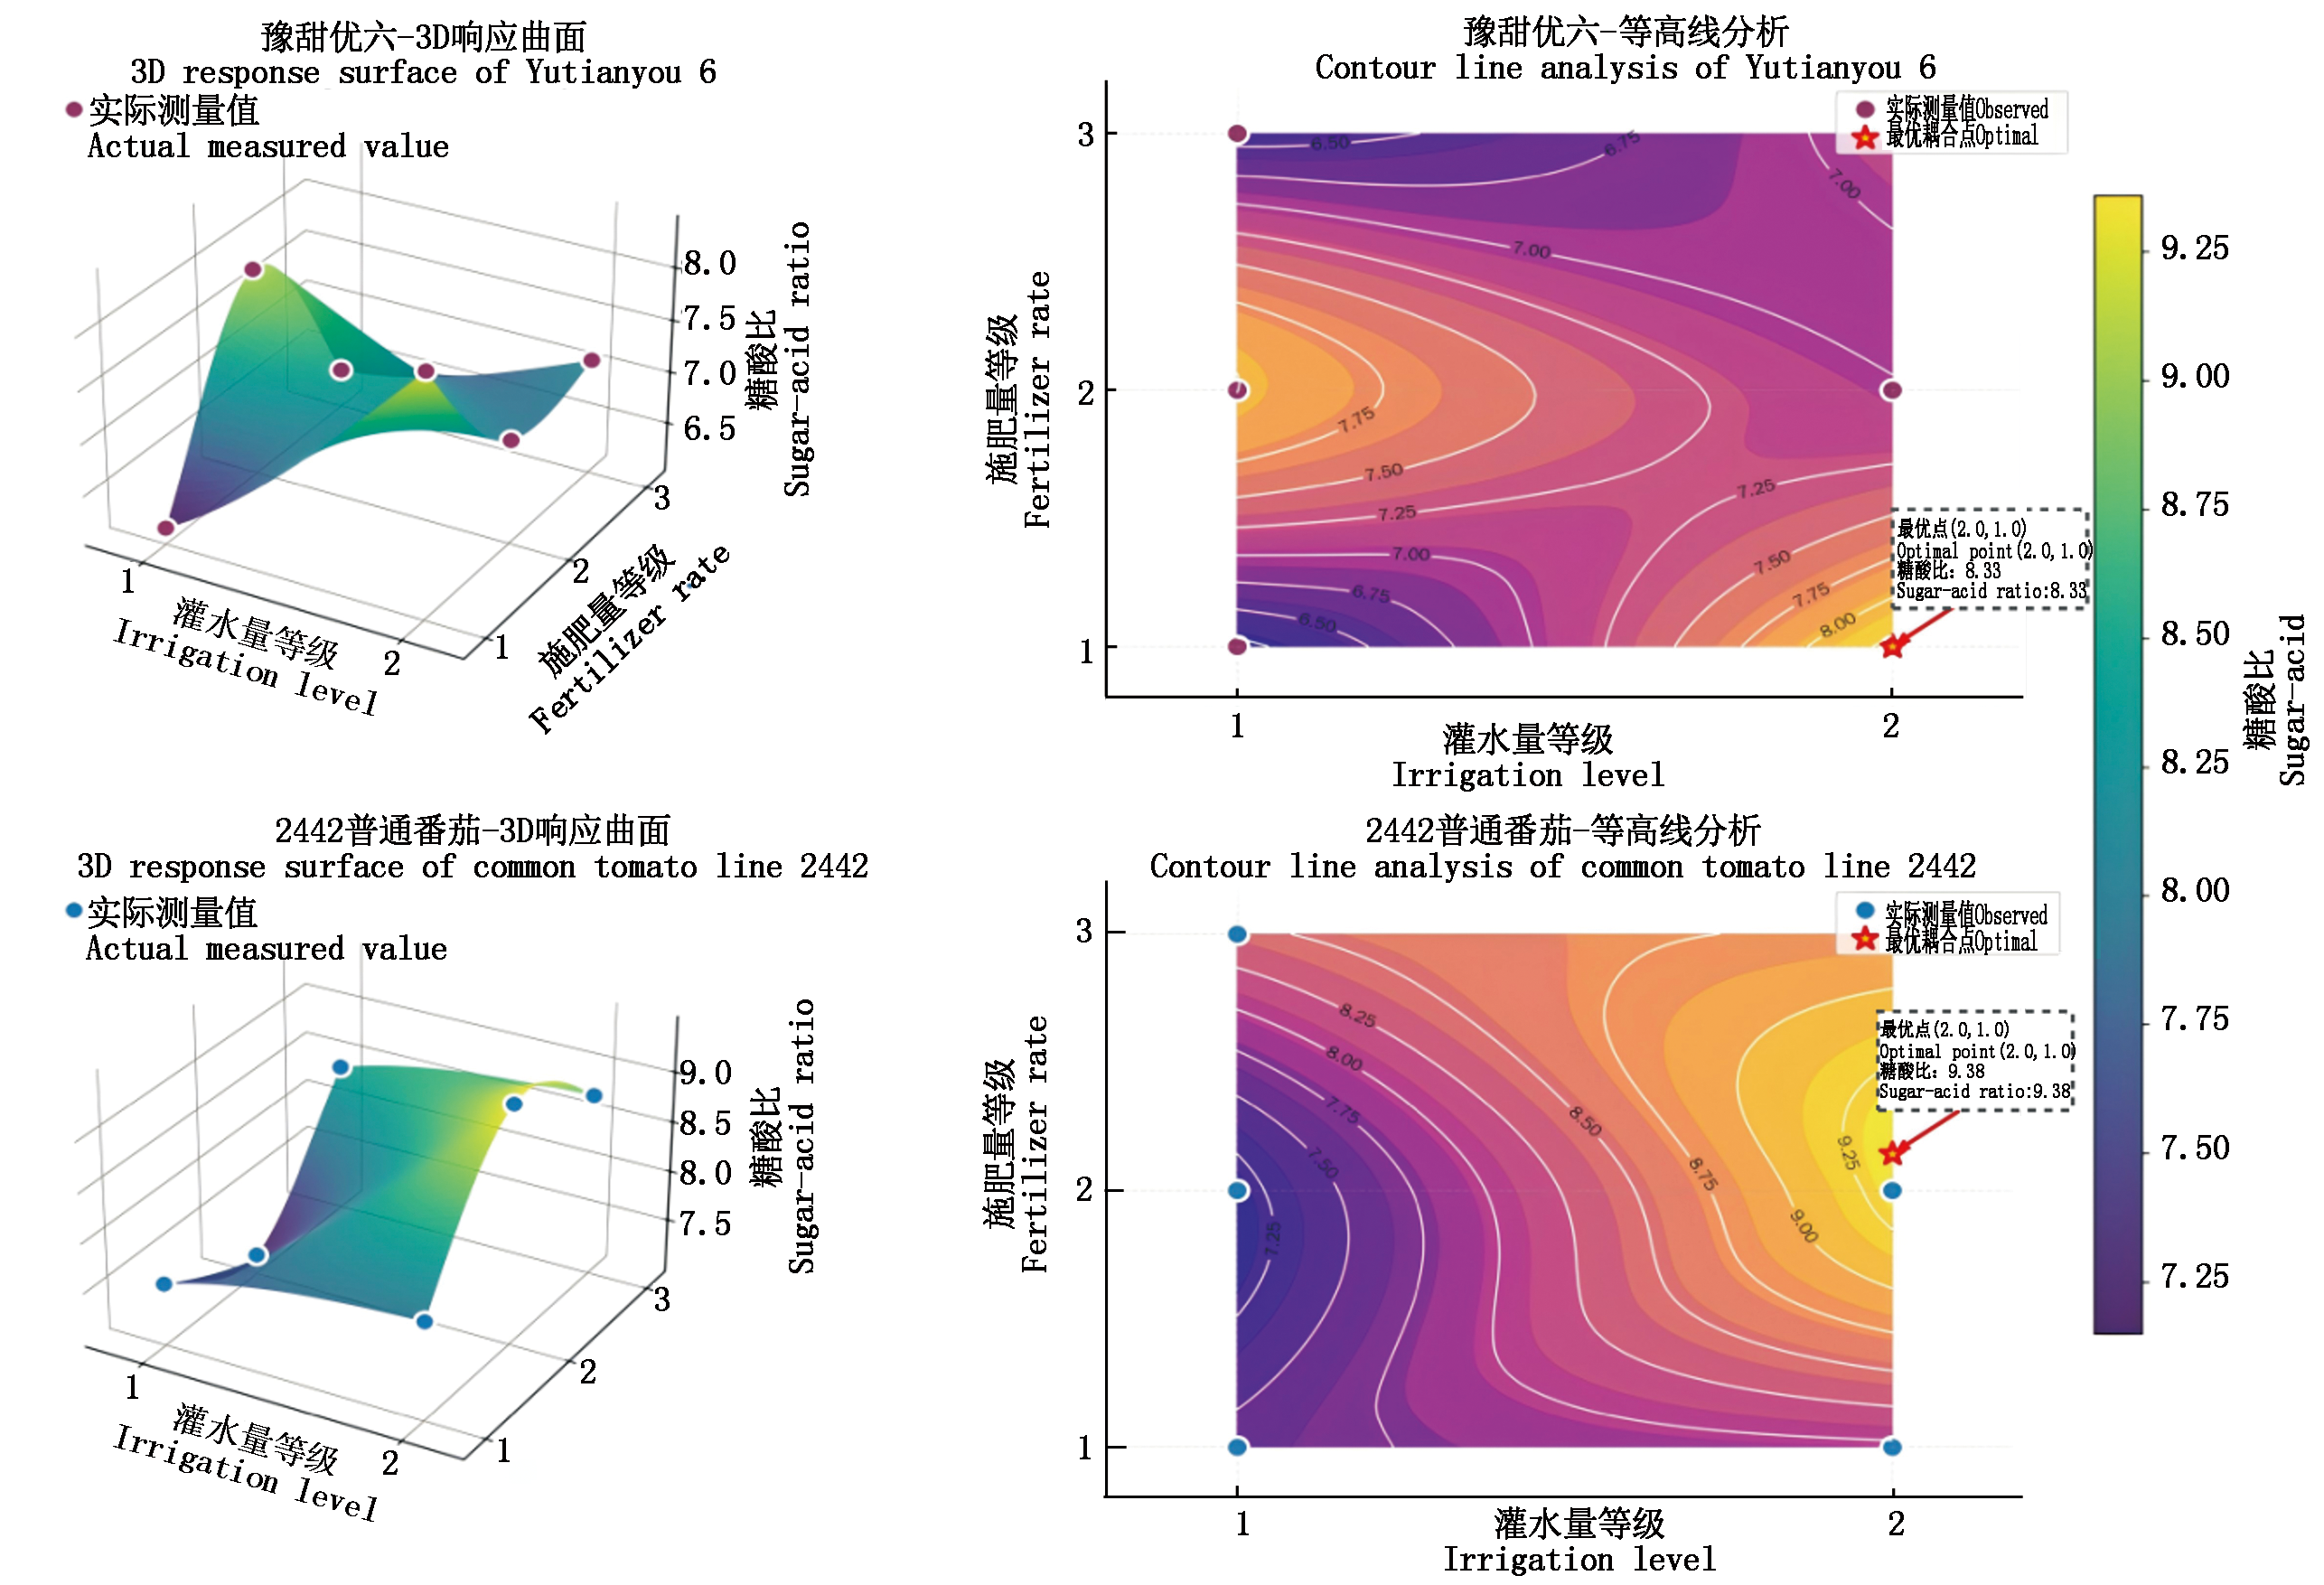
This study investigated the function of the cotton gene GhMYB113 in resistance to Verticillium wilt.GhMYB113 encoded a 249 amino acid protein with a molecular weight of 29.07 ku and an isoelectric point of 6.46.Phylogenetic and subcellular localization analyses revealed that GhMYB113 shared the highest homology with soybean GmMYB113 and was localized to the nucleus.Expression profiling by qPCR showed that GhMYB113 was predominantly expressed in roots and was upregulated upon infection by Verticillium dahliae as well as by treatment with jasmonic acid(JA)and salicylic acid(SA),suggesting a role in disease resistance signaling.Silencing of GhMYB113 via virus-induced gene silencing(VIGS)led to a significant decrease in leaf anthocyanin accumulation and downregulated the expression of key anthocyanin biosynthesis genes,indicating that GhMYB113 positively regulates anthocyanin synthesis in cotton.This was further confirmed using a GUS reporter system,which demonstrated that GhMYB113 activates the promoter of GhDFR.Inoculation assays performed on GhMYB113-silenced plants resulted in more severe disease symptoms,higher disease index(DI)values,increased disease incidence,enhanced stem browning,and significantly greater fungal biomass in stems compared with control plants,demonstrating that GhMYB113 positively regulates cotton resistance to Verticillium wilt.In summary,GhMYB113 plays a critical role in both anthocyanin biosynthesis and Verticillium wilt resistance in cotton,providing a theoretical foundation for breeding disease-resistant cotton varieties.
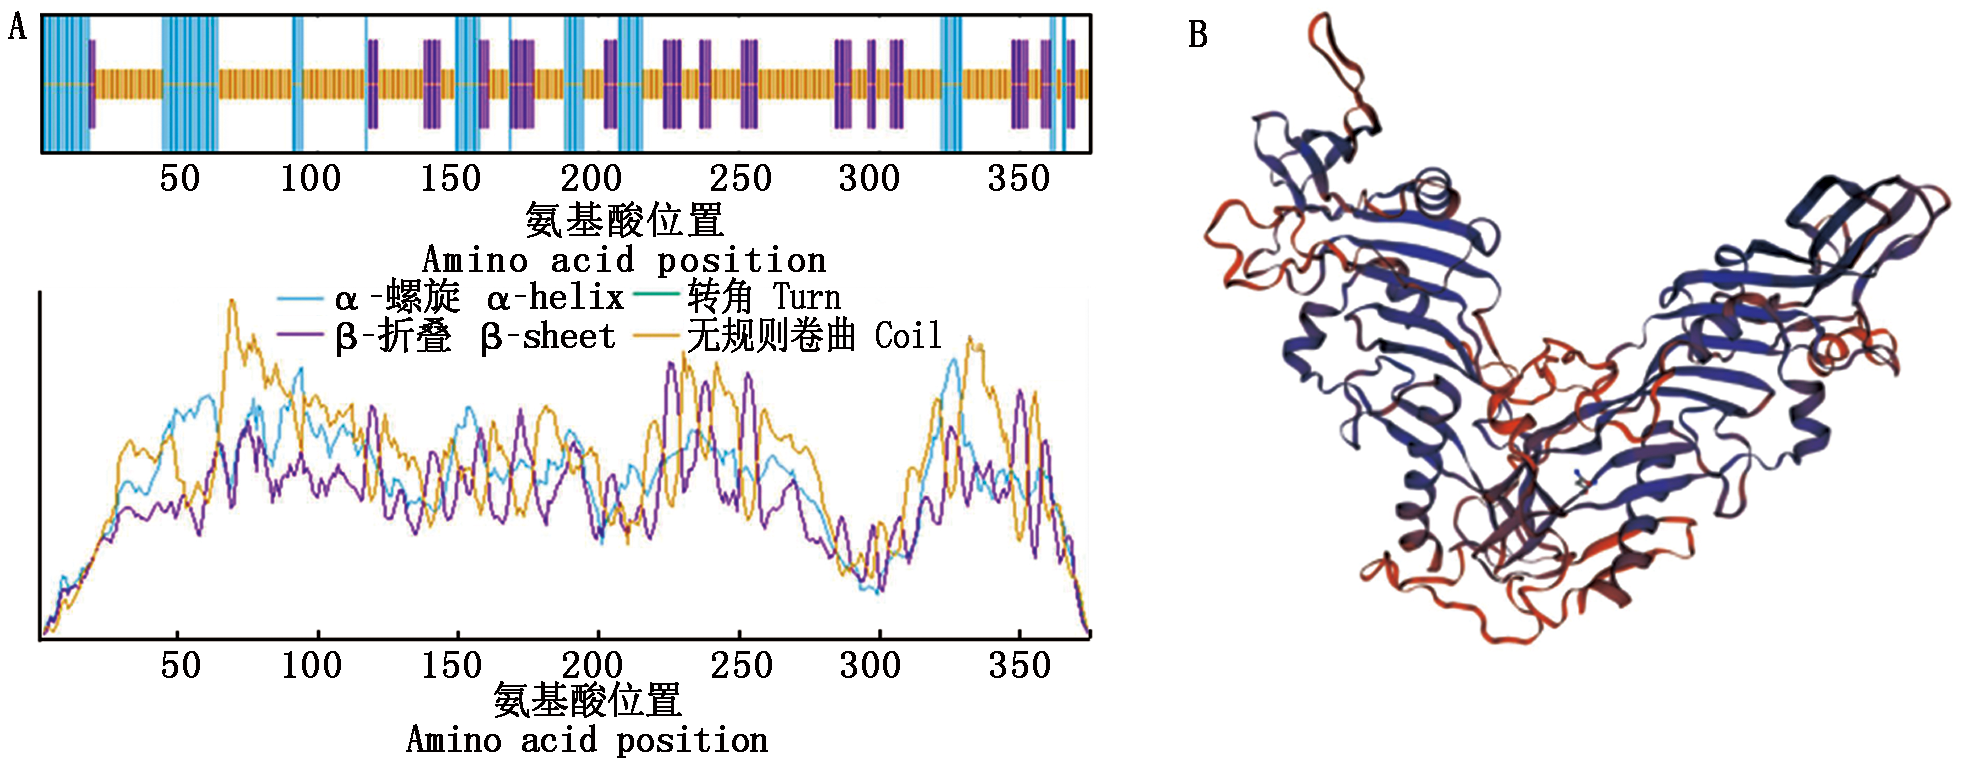
In order to explore the gene function of nerve growth factor(NGF),and to clarify the expression differences of NGF in different tissues of Red Steppe cattle and its influence on the myogenic differentiation of bovine skeletal muscle satellite cells(bSMSCs),collect various tissues of the Red Steppe cattle,clone and obtain the complete CDS region of the NGF gene,and then conduct bioinformatics and tissue expression analysis on it.bSMSCs were isolated from the deep tissues of the hind limbs of neonatal calves.The expression of Pax7,Desmin and MyoD1,the specific markers of bSMSCs,was detected by immunofluorescence staining.In vitro,2% horse serum was used to induce myogenic differentiation of bSMSCs by serum deprivation method,and the expression patterns of NGF mRNA and protein at different stages of differentiation were detected.The results showed that the length of the CDS region of bovine NGF gene was 726 bp,encoding 241 amino acids.The amino acid sequence of Red Steppe NGF gene is located in a branch of house mouse,red deer and sheep,with high homology and close kinship.The molecular weight of NGF protein was about 26.67 ku,the molecular formula was C1179H1861N349O339S10,the theoretical isoelectric point was 9.72,the half-life was 30 h,and the NGF protein was water soluble.NGF protein had signal peptide and was secreted protein.There were 42 potential phosphorylation sites.There were three potential sites for N-glycosylation.The secondary structure of NGF protein was composed of α-helix,β-sheet,β-turn and random coil,and the results of tertiary structure prediction were consistent with those of NGF protein.There were significant differences in the expression level of NGF gene in different tissues,with the highest expression level in liver,followed by kidney and tumour.The expression of the NGF gene was continuously upregulated during the myogenic differentiation process of bSMSCs,and there was a significant difference on the 7th day.This study successfully cloned the CDS region sequence of the NGF gene in the grassland,and established the tissue expression profile of the NGF gene in the Red Steppe cattle.The expression pattern of the NGF gene during the myogenic differentiation process of bSMSCs was clarified.

To clone the full-length cDNA sequence of the MSTN(Myostatin)gene from Gymnocypris przewalskii,clarify its expression characteristics in different tissues of G.przewalskii and expression differences in the muscle tissues of different Cyprinidae fishes,and provide basic data for exploring the molecular mechanism underlying the slow growth of G.przewalskii.5'RACE and 3'RACE techniques were used to clone the full-length cDNA sequence of the MSTN gene from G.przewalskii.Bioinformatics methods were employed to analyze its sequence characteristics,and Quantitative Real-time PCR(RT-qPCR)was used to detect the differential expression of this gene in different tissues of G.przewalskii and muscle tissues of different Cyprinidae fishes.The results showed that the full-length cDNA sequence of the MSTN gene from G.przewalskii was 2 168 bp,including an open reading frame(ORF)of 1 128 bp encoding 375 amino acids,with ATG as the start codon and TGA as the stop codon.The predicted encoded protein contained 70 threonine(Thr)phosphorylation sites.Phylogenetic analysis showed that the MSTN gene of G.przewalskii clustered with homologous genes of other Cyprinidae fishes,indicating a close genetic relationship.RT-qPCR results revealed that MSTN was expressed in gill,kidney,liver,heart,brain,muscle,and skin tissues of G.przewalskii,with relatively high expression levels in the brain and muscle tissues.Among different Cyprinidae fishes,the expression level of the MSTN gene was relatively higher in G.przewalskii and Gymnocypris eckloni,while relatively lower in Ctenopharyngodon idella,Carassius auratus,and Megalobrama amblycephala.The full-length cDNA sequence of the MSTN gene from G.przewalskii was successfully cloned,and its sequence characteristics and evolutionary status were clarified,the expression differences of this gene in different tissues of G.przewalskii and among different Cyprinidae fishes were revealed.
Bimonthly, Started in 1962
CN 13-1101/S
ISSN 1000-7091
CODEN: HHUOA6
Responsible Institution: Hebei Academy of Agriculture and Forestry Sciences
Sponsored by: the Academy of Agricultural Sciences and Agricultural Association of Hebei, Beijing, Tianjin, Shanxi, Henan and Inner Mongolia.
Editor-in-chief: Qiang Zhang
Edited and Published by: Editorial Department of Acta Agriculturae Boreali-Sinica
《Acta Agriculturae Boreali-Sinica》Official Website
Wechat Official Account

Copyright © 2022 《Acta Agriculturae Boreali-Sinica》

